Skip to content

Food and Drug Regulations

C.R.C., c. 870

In forceCurrent to September 30, 2026 HistoryMade under: Food and Drugs Act

Loading…

Part C — Drugs › Division 1

General

Amended on June 17, 2026

  •  (1) In this Part

    acetaminophen product

    acetaminophen product has the same meaning as in Division 9; (produit d’acétaminophène)

    adult standard dosage unit

    adult standard dosage unit has, with reference to a drug, the same meaning as in Division 9; (dose normale pour adultes)

    adverse drug reaction

    adverse drug reaction means a noxious and unintended response to a drug, which occurs at doses normally used or tested for the diagnosis, treatment or prevention of a disease or the modification of an organic function; (réaction indésirable à une drogue)

    antibiotic

    antibiotic means any drug or combination of drugs such as those named in C.01.410 to C.01.592 which is prepared from certain micro-organisms, or which formerly was prepared from micro-organisms but is now made synthetically and which possesses inhibitory action on the growth of other micro-organisms; (antibiotique)

    authorization holder

    authorization holder[Repealed, SOR/2017-259, s. 1]

    brand name

    brand name means, with reference to a drug, the name, whether or not including the name of any manufacturer, corporation, partnership or individual, in English or French,

    • (a) that is assigned to the drug by its manufacturer,

    • (b) under which the drug is sold or advertised, and

    • (c) that is used to distinguish the drug; (marque nominative)

    case report

    case report means a detailed record of all relevant data associated with the use of a drug in a subject; (fiche d’observation)

    children’s standard dosage unit

    children’s standard dosage unit has, with reference to a drug, the same meaning as in Division 9; (dose normale pour enfants)

    child resistant package

    child resistant package means a package that meets the requirements of subsection (2); (emballage protège-enfants)

    common name

    common name means, with reference to a drug, the name in English or French by which the drug is

    • (a) commonly known, and

    • (b) designated in scientific or technical journals, other than the publications referred to in Schedule B to the Act; (nom usuel)

    COVID-19

    COVID-19 means the coronavirus disease 2019; (COVID-19)

    COVID-19 drug

    COVID-19 drug means a drug, other than a veterinary health product, that is manufactured, sold or represented for use in relation to COVID-19; (drogue contre la COVID-19)

    discontinue

    discontinue means, in respect of the sale of a drug by the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for the drug, to permanently cease the sale of the drug in Canada; (cesser)

    expiration date

    expiration date means

    • (a) in the case of a drug in dosage form, the earlier of the following dates, expressed at minimum as a year and month:

      • (i) the date up to and including which the drug maintains its labelled potency, purity and physical characteristics, and

      • (ii) the date after which the manufacturer recommends that the drug not be used; and

    • (b) in the case of an active ingredient, whichever of the following dates is applicable, expressed at minimum as a year and month:

      • (i) the retest date, or

      • (ii) the date after which the manufacturer recommends that the active ingredient not be used; (date limite d’utilisation)

    flavour

    flavour means a non-medicinal ingredient or combination of non-medicinal ingredients added to a drug solely to produce or mask a particular taste. It does not include an ingredient or combination of ingredients that impart only a sweet taste to the drug; (saveur)

    fragrance

    fragrance means a non-medicinal ingredient or combination of non-medicinal ingredients added to a drug to produce or mask a particular odour; (parfum)

    immediate container

    immediate container means the receptacle that is in direct contact with a drug; (récipient immédiat)

    internal use

    internal use means ingestion by mouth or application for systemic effect to any part of the body in which the drug comes into contact with mucous membrane; (usage interne)

    ISAD Interim Order

    ISAD Interim Order means the Interim Order Respecting the Importation, Sale and Advertising of Drugs for Use in Relation to COVID-19 made by the Minister on September 16, 2020 and published in Part I of the Canada Gazette on October 3, 2020; (arrêté d’urgence IVPD)

    List A

    List A means the document, entitled List of Certain Antimicrobial Active Pharmaceutical Ingredients, that is published by the Government of Canada on its website, as amended from time to time; (Liste A)

    List B

    List B means the document, entitled List of Certain Veterinary Drugs Which May Be Imported But Not Sold, that is published by the Government of Canada on its website, as amended from time to time; (Liste B)

    List C

    List C means the document, entitled List of Veterinary Health Products, that is published by the Government of Canada on its website, as amended from time to time; (Liste C)

    List D

    List D means the document entitled List of Certain Non-prescription Drugs for Distribution as Samples that is published by the Government of Canada on its website, as amended from time to time; (Liste D)

    List of Drugs for the Purposes of Paragraphs (b) and (c) of the Definition drug in Subsection C.01.014.8(1)

    List of Drugs for the Purposes of Paragraphs (b) and (c) of the Definition drug in Subsection C.01.014.8(1) means the List of Drugs for the Purposes of Paragraphs (b) and (c) of the Definition drug in Subsection C.01.014.8(1) of the Food and Drug Regulations that is published by the Government of Canada on its website, as amended from time to time; (Liste de drogues pour l’application des alinéas b) et c) de la définition de drogue au paragraphe C.01.014.8(1))

    non-medicinal ingredient

    non-medicinal ingredient means a substance — other than the pharmacologically active drug — that is added during the manufacturing process and that is present in the finished drug product; (ingrédient non médicinal)

    official drug

    official drug means any drug

    • (a) for which a standard is provided in these Regulations, or

    • (b) for which no standard is provided in these Regulations but for which a standard is provided in any of the publications mentioned in Schedule B to the Act; (drogue officielle)

    parenteral use

    parenteral use means administration of a drug by means of a hypodermic syringe, needle or other instrument through or into the skin or mucous membrane; (usage parentéral)

    per cent

    per cent means per cent by weight unless otherwise stated; (pour cent)

    pharmaceutical ink

    pharmaceutical ink means a non-medicinal ingredient or combination of non-medicinal ingredients used to imprint the drug with marks or symbols; (encre pharmaceutique)

    pharmacist

    pharmacist means a person who

    • (a) is registered or otherwise entitled under the laws of a province to practise pharmacy, and

    • (b) is practising pharmacy in that province; (pharmacien)

    pharmacy technician

    pharmacy technician means a person who

    • (a) is registered or otherwise entitled under the laws of a province to practise as a pharmacy technician; and

    • (b) is practising as a pharmacy technician in that province; (technicien en pharmacie)

    practitioner

    practitioner means a person who

    • (a) is entitled under the laws of a province to treat patients with a prescription drug, and

    • (b) is practising their profession in that province; (praticien)

    prescription

    prescription means an order given by a practitioner directing that a stated amount of any drug or mixture of drugs specified therein be dispensed for the person named in the order; (ordonnance)

    proper name

    proper name means, with reference to a drug, the name in English or French

    • (a) assigned to the drug in section C.01.002,

    • (b) that appears in bold-face type for the drug in these Regulations and, where the drug is dispensed in a form other than that described in this Part, the name of the dispensing form,

    • (c) specified in the Canadian licence in the case of drugs included in Schedule C or Schedule D to the Act, or

    • (d) assigned in any of the publications mentioned in Schedule B to the Act in the case of drugs not included in paragraph (a), (b) or (c); (nom propre)

    salicylate product

    salicylate product has the same meaning as in Division 9; (produit de salicylate)

    serious adverse drug reaction

    serious adverse drug reaction means a noxious and unintended response to a drug that occurs at any dose and that requires in-patient hospitalization or prolongation of existing hospitalization, causes congenital malformation, results in persistent or significant disability or incapacity, is life-threatening or results in death; (réaction indésirable grave à une drogue)

    serious unexpected adverse drug reaction

    serious unexpected adverse drug reaction means a serious adverse drug reaction that is not identified in nature, severity or frequency in the risk information set out on the label of the drug; (réaction indésirable grave et imprévue à une drogue)

    shortage

    shortage means, in respect of a drug, a situation in which the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for the drug is unable to meet the demand for the drug in Canada; (pénurie)

    teaspoon

    teaspoon means, for the purpose of calculation of dosage, a volume of 5 cubic centimetres; (cuillerée à thé)

    test group

    test group means a group that meets the requirements of subsection (3); (groupe d’essai)

    veterinary health product

    veterinary health product means any of the following drugs that is in dosage form and that is not manufactured, sold or represented for use in the diagnosis, treatment, mitigation or prevention of a disease, disorder or abnormal physical state, or its symptoms:

    • (a) a substance set out in Column I of Part 1 of List C that is consistent with the descriptive information set out in Columns II to V, or any combination of any substances in which all the medicinal ingredients are substances set out in Column I of Part 1 of that list if that combination is, in respect of each of those substances, consistent with the descriptive information set out in Columns II and III and the descriptive information set out in Columns IV and V that is, within each of those columns, common to those substances;

    • (b) a homeopathic medicine set out in Column I of Part 2 of List C that is consistent with the descriptive information set out in Columns II to V, or any combination of homeopathic medicines set out in Column I of Part 2 of that list if that combination is, in respect of each of those homeopathic medicines, consistent with the descriptive information set out in Columns II and III and the descriptive information set out in Columns IV and V that is, within each of those columns, common to those homeopathic medicines; and

    • (c) a traditional medicine set out in Column I of Part 3 of List C that is consistent with the descriptive information set out in Columns II to V, or any combination of traditional medicines set out in Column I of Part 3 of that list if that combination is, in respect of each of those traditional medicines, consistent with the descriptive information set out in Columns II and III and the descriptive information set out in Columns IV and V that is, within each of those columns, common to those traditional medicines; (produit de santé animale)

    withdrawal period

    withdrawal period means the length of time between the last administration of a drug to an animal and the time when tissues or products collected from the treated animal for consumption as food contain a level of residue of the drug that would not likely cause risk to human health. (délai d’attente)

  • (1.1) For the purposes of the Act, serious adverse drug reaction has the same meaning as in subsection (1).

  • (2) A child resistant package is a package that

    • (a) when tested in accordance with an acceptable method,

      • (i) in the case of a test group comprising children, cannot be opened

        • (A) by at least 85 per cent of those children prior to a demonstration to them of the proper means of opening the package, and

        • (B) by at least 80 per cent of those children after the demonstration, and

      • (ii) in the case of a test group comprising adults

        • (A) can be opened by at least 90 per cent of those adults, and

        • (B) where the package is designed so that, once opened and reclosed, it continues to meet the requirements of subparagraph (i), can be so reclosed by at least 90 per cent of those adults; or

    • (b) complies with the requirements of one of the following standards, namely,

      • (i) Canadian Standards Association Standard CAN/CSA-Z76.1-M90, entitled Recloseable Child-Resistant Packages, published January 1990, as amended from time to time,

      • (ii) European Standard EN 28317:1992, entitled Child-resistant packaging — Requirements and testing procedures for reclosable packages, as adopted by the European Committee for Standardization on October 30, 1992, recognized by the British Standards Institution, and effective February 15, 1993 and by the Association française de normalisation, and effective December 20, 1992, and which reiterates fully the international standard ISO 8317:1989, as amended from time to time, and

      • (iii) Code of Federal Regulations (United States), Title 16, Section 1700.15, entitled Poison prevention packaging standards, as amended from time to time.

  • (3) For the purposes of this section, test group means

    • (a) in relation to children, a group of at least 200 children who

      • (i) are healthy and have no obvious physical or mental disability,

      • (ii) are between 42 and 51 months of age, and

      • (iii) represent evenly, within plus or minus 10 per cent, each monthly age between 42 and 51 months calculated to the nearest month; and

    • (b) in relation to adults, a group of at least 100 adults who

      • (i) are healthy and have no obvious physical or mental disability,

      • (ii) are between 18 and 45 years of age, and

      • (iii) represent evenly, within plus or minus 10 per cent, each yearly age between 18 and 45 years calculated to the nearest year.

  • (4) For the purpose of this section, an amendment from time to time to a standard referred to in paragraph (2)(b) becomes effective 18 months after the date designated by the competent authority as the effective date for the amendment.

  • SOR/80-543, s. 1
  • SOR/85-966, s. 1
  • SOR/86-93, s. 1
  • SOR/87-484, s. 1
  • SOR/92-654, s. 1
  • SOR/93-202, s. 1
  • SOR/95-411, s. 1
  • SOR/95-521, s. 1
  • SOR/96-399, s. 1
  • SOR/96-240, s. 1
  • SOR/97-543, s. 5
  • SOR/2010-105, s. 1
  • SOR/2013-74, s. 1
  • SOR/2013-122, s. 3
  • SOR/2016-139, s. 1
  • SOR/2017-76, s. 1
  • SOR/2017-133, s. 1
  • SOR/2017-259, s. 1
  • SOR/2020-74, s. 1
  • SOR/2021-45, s. 1
  • SOR/2021-46, s. 11(E)
  • SOR/2022-197, s. 8
  • SOR/2023-247, s. 1(F)
  • SOR/2026-96, s. 1
.

 The Proper Name of a drug shown opposite an item number in the following Table in the column headed “Chemical Names and Synonyms” shall be the name shown opposite that item number in the column headed “Proper Names”.

TABLE

Item No.Proper NamesChemical Names and Synonyms
A.1Acepromazine 2-acetyl-10-(3-dimethylaminopropyl) iephenothiazine
A.2Acetaminophen p-Acetaminophenol, Paracetamol, p-Hydroxyacetanilide: N-acetyl-p-aminophenol
A.3Acetanilide: Acetanilid Acetylaminobenzene: Antifebrin: Phenylacetamide
A.4Acetylsalicylic Acid Acetylsalicylic acid
A.5Allopurinol 1-H-Pyrazolo [3,4-d] pyrimidin-4-ol: 4-Hydroxypyrazolo (3,4-d) pyrimidine
A.6Amantadine 1-Adamantanamine
A.7Aminocaproic acid 6-Aminohexanoic acid
A.8Aminopterin N-[4-(2,4-diamino-6-pteridyl methyl) amino-benzoyl]-L- glumatic acid
A.9Aminopyrine: Amidopyrine 1,5-dimethyl-2-phenyl-4-dimethylamino-3-pyrazolone: Dimethylaminophenazone
A.10Amitriptyline 3-(3-Dimethylaminopropylidene)-1,2: 4,5-dibenzocyclohepta-1,4-diene
A.11Azacyclonol α,α-diphenyl-4-piperidinecarbinol
B.1Bemegride 3-Ethyl-3-methylglutarimide
B.2Benactyzine Dimethylaminoethyl-1,1-diphenylglycolate
B.3Bendroflumethiazide 3-benzyl-3,4-dihydro-6-(trifluoro- methyl)-2H-1,2,4-benzothiadiazine-7-sulfonamide-1,1-dioxide: Bendrofluazide (B.A.N.)
B.4Betahistine 2-[2-(Methylamino)ethyl] pyridine
B.5Bethanidine N-Benzyl-N′N″-dimethylguanidine: 1-Benzyl-2,3-dimethylguanidine
B.6Bretylium tosylate N-2-Bromobenzyl-N-ethyl-N, N-dimethylammonium tosylate (Tosylic acid is trivial name for p-toluenesulphonic acid)
B.7Bromisoval 2-monobromoisovalerylurea: Bromisovalum: Bromvalitone
C.1Calcium Carbimide Calcium cyanamide
C.2Captodiamine 4-butylthio-α-phenylbenzyl-2- dimethylaminoethylsulfide
C.3Carisoprodol N-Isopropyl-2-methyl-2-propyl-1, 3-propanediol dicarbamate
C.4Carphenazine 1-[10-(3[4-(2-Hydroxyethyl)-1-piperazinyl]propyl) phenothiazin-2yl]-1-propapone
C.5Cephaloridine 7-[(2-Thienyl) acetamido]-3-(1-pyridylmethyl)-3-cephem-4-carboxylic acid betaine
C.6Chlormezanone 2-(4-chlorophenyl)-3-methyl-4-methathiazanone-1,1-dioxide: Chlormethazone: Chlormethazanone
C.7Chloromethapyrilene N,N-dimethyl-N′-(2-pyridyl)-N′-(5-chloro-2-thenyl)-ethylenediamine: Chlorothen
C.8Chlorphentermine 4-Chloro-α,α-dimethylphenethylamine
C.9Cinchocaine 2-butoxy-N-(2-diethylaminoethyl) cinchoninamide: Dibucaine
C.10Cinchophen 2-phenylquinoline-4-carboxylic acid: Quinophan
C.11Clofibrate Ethyl 2-(p-chlorophenoxy)-2-methylpropionate
C.12Clomiphene 1-Chloro-2-[4-(2-diethylamino-ethoxy)phenyl]-1,2-diphenylethylene: 2-[p-(2-Chloro-1,2-diphenylvinyl)phenoxy] triethylamine
D.1Desipramine 5-(3-Methylaminopropyl)-10,11-dihydro-5H-dibenz[b,f]azepine
D.2Diazepam 7-Chloro-1,3-dihydro-1-methyl-5-phenyl-2H-1,4-benzodiazepin-2-one
D.3Diethylpropion 1-phenyl-2-diethylaminopropanone-1
D.4Diphenidol 1,1-Diphenyl-4-piperidinobutan-1-ol
D.5Disulfiram Tetraethylthiuram disulphide
E.1Ectylurea 2-ethyl-cis-crotonylurea
E.2Emylcamate 1-Ethyl-1-methylpropyl carbamate
E.3Ethacrynic Acid [2,3-Dichloro-4-(2-methylenebutyryl) phenoxy] acetic acid: 2,3-Dichloro-4-(2-ethylacryloyl) phenoxyacetic acid
E.4Ethchlorvynol 3-(2-chlorovinyl)-1-pentyn-3-ol
E.5Ethinamate 1-ethynylcyclohexyl carbamate
E.6Ethionamide 2-Ethylisonicotinthioamide
E.7Ethomoxane 2-n-Butylaminomethyl-8-ethoxy-benzo-1,4-dioxan
E.8Ethyl Trichloramate Ethyl n-[1-(2,2,2,-trichloro-1-hydroxyethyl)] carbamate
E.9Etryptamine 3-(2-Aminobutyl) indole
E.10Etymemazine 10-(3-Dimethylamino-2-methylpro-pyl)-2-ethylphenothiazine
F.1Fluphenazine 10-{3-[4-(2-Hydroxyethyl) piperazin-1-yl] propyl}-2-tri-fluoromethylphenothiazine
F.2Furosemide 4-Chloro-N-furfuryl-5-sulphamoylanthranilic acid: Frusemide (B.A.N.)
G.1Glyburide 5-chloro-N-[2-[4-[[[(cyclohexylamino carbonyl]amino]sulfonyl]phenyl]ethyl]-2-methoxy benzamide: 1-4[4-[2-(5-chloro-2-methoxybenzamido)ethyl]phenyl- sulfonyl]-3-cyclohexylurea: Glibenclamide
H.1Haloperidol 4-(4-Chlorophenyl)-1-[3-(4-fluorobenzoyl) propyl]-piperidin-4-ol: 4-[4-(p-Chlorophenyl)-4-hydro-xypiperidino]-4’-fluorobutyro-phenone
H.2Hydroxychloroquine 7-Chloro-4[4-(N-ethyl-N-2-hydro-xyethylamino)-1-methylbutyl-amino] quinoline
H.3Hydroxyzine 1-(p-chloro-α-phenylbenzyl)-4-(2-hydroxy ethoxyethyl) piperazine
I.1Idoxuridine 5-Iodo-2′-deoxyuridine
I.2Imipramine 5-(3-dimethylaminopropyl)-10,11-dihydro-5H-dibenz[b,f]azepine
I.3Indomethacin 1-(p-Chlorobenzoyl)-5-methoxy-2-methyl-indole-3-acetic acid
I.4Iproniazid 1-isonicotinoyl-2-isopropylhydrazine
I.5Isocarboxazid 3-N-Benzylhydrazinocarbonyl-5-methylisoxazole
I.6Isoproterenol 3,4-Dihydroxy-α-[isopropylamino) methyl] benzyl alcohol: Isoprenaline
L.1Liothyronine L-α-Amino-3-[(4-hydroxy-3-iodophenoxy)-3,5-di-iodo-phenyl] propionic acid
M.1Mefenamic acid N-(2,3-Xylyl)-anthranilic acid
M.2Melphalan 4-Di-(2-chlorethyl)amino-L-phenylalanine
M.3Mepazine 10-[(1-methyl-3-piperidyl) methyl] phenothiazine
M.4Mephenesin 3-o-toloxy-1,2-propanediol
M.5Mephenoxalone 5-(o-Methoxyphenoxymethyl)-2-oxazolidinone
M.6Meprobamate 2,2-di(carbamoylmethyl) pentane
M.7Methaqualone 2-Methyl-3-o-tolyquinazolin-4-one: 2-Methyl-3-o-tolyl-4-quinazolone
M.8Methisazone 1-Methylindoline-2,3-dione-3-thiosemicarbazone: N-Methylisatin-ß-thiosemicarbazone
M.9Methotrimeprazine 10-[3-(2-Methyl)dimethylamino propyl]-2-methoxyphenothiazine: Levomepromazine
M.10Methyldopa 1-3(3,4-Dihydroxyphenyl)-2-methylalanine
M.11Methylparafynol 3-methyl-1-pentyn-3-ol: Methylpentynol
M.12Methylphenidate Methyl-1-phenyl-1-(2-piperidyl) acetate
M.13Methyprylon 3,3-diethyl-5-methyl-2,4-piperidinedione
M.14Methysergide 1-(Hydroxymethyl)propylamide of 1-methyl-d-lysergic acid
M.15Metyrapone 2-Methyl-1,2-di(3-pyridyl)propan-1-one
N.1Nalidixic Acid 1-Ethyl-7-methyl-4-oxo-1,8-naphthyridine-3-carboxylic acid
N.2Nialamide 1-[2-(benzycarbamyl)ethyl]-2-isonicotinoyl-hydrazine
N.3Nortriptyline 3-(3-Methylaminopropylidene)-1,2, 4,5-dibenzocyclohepta-1,4-diene
O.1Oxanimide 2-ethyl-3-propyl-glycidamide
O.2Oxazepam 7-Chloro-1,3-dihydro-3-hydroxy-5-phenyl-1,4-benzodiazepin-2-one
O.3Oxyphenbutazone 4-n-Butyl-2-(4-hydroxyphenyl)-1-phenyl-pyrazolidine-3,5-dione
P.1Paramethadione 3,5-dimethyl-5-ethyl-2,4-oxazolidinedione
P.2Pargyline N-Benzyl-N-methylprop-2-ynylamine
P.3Pemoline 2-Imino-5-phenyloxazolidin-4-one
P.4Pentazocine 1,2,3,4,5,6-Hexahydro-8-hydroxy-6,11-dimethyl-3-(3-methylbut-2-enyl)-2,6-methano-3-benzazocine: 1,2,3,4,5,6-Hexahydro-6,11-dimethyl-3-(3-methyl-2-butenyl)-2,6-methano-3-benzazocin-8-ol
P.5Pentolinium Tartrate NN′-Pentamethylenedi-(methylpryrrolidinium hydrogen, tartrate)
P.6Perphenazine 2-chloro-10-{3-[1-(2-hydroxyethyl)-4-piperazinyl]propyl} phenothiazine
P.7Phacetoperane l-1-Phenyl-1(2′-piperidyl)-1-acetoxymethane
P.8Phenacemide (Phenylacetyl)urea
P.9Phenacetin p-acetphenetidin: Acetphenetidin: Acetophenetidin: p-ethoxyacetanilid
P.10Phenaglycodol 2-p-chlorophenyl-3-methyl-2,3-butanediol
P.11Phendimetrazine 3,4-Dimethyl-2 Phenylmorpholine
P.12Phenelzine 2-phenylethylhydrazine
P.13Phenformin N′-ß-phenethylformamidinyliminourea
P.14Pheniprazine α-Methylphenethylhydrazine
P.15Phenmetrazine Tetrahydro-3-methyl-2-phenyl-1,4-oxazine: 3-methyl-2-phenylmorpholine
P.16Phentermine α, α-Dimethylphenethylamine: phenyl-tert-butylamine
P.17Phenylindanedione 2-phenylindane-1,3-dione
P.18Phenyltoloxamine N,N-dimethyl-2-(α-phenyl-o-tolyloxy) ethylamine
P.19Pholedrine p-(4-hydroxyphenyl)-isopropylmethylamine
P.20Piperliate 1-piperidine-ethanol benzilate
P.21Pipradol Diphenyl-2-piperidylmethanol
P.22Prochlorperazine 2-Chloro-10-[3-(1-methyl-4-piperazinyl) propyl]phenothiazine
P.23Prodilidine 1,2-Dimethyl-3-phenyl-3-pyrrolidinyl propionate
P.24Propranolol 1-(Isopropylamino)-3-(1-naphthyloxy)-2-propanol
P.25Prothipendyl 9-(3-Dimethylaminopropyl)-10-thia-1,9-diaza-anthracene
P.26Protriptyline 7-(3-Methylaminopropyl)-1,2:5,6-dibenzocycloheptatrien: N-Methyl-5H-dibenzo [a, d] cycloheptene-5-propylamine
P.27Pyrazinamide Pyrazinoic acid amide
R.1Rifampin 3-{[(4-methyl-1-piperazinyl)imino]methyl} rifamycin SV : Rifampicin (I.N.N.) (Rifamycin SV is an antibiotic produced by Streptomyces mediterranei)
S.01Sodium Cromoglycate 4H-1-Benzopyran-2-carboxylic acid, 5,5′-[(2-hydroxy-1,3-propanediyl) bis(oxy)]bis[4-oxo-,disodium salt]:
Disodium 5,5′-(2-hydroxytrimethylenedioxy) bis[4-oxo-4H-1-benzopyran-2- carboxylate]: Disodium 4,4′-dioxo-5,5′-(2-hydroxytrimethylenedioxy)di (chromene-2-carboxylate): Cromolyn Sodium (USP): Disodium Cromoglycate
S.1Sulfameter 2-(4-Aminobenzenesulphonamido)-5-methoxypyrimidine: N′-(5-methoxy-2-pyrimidinyl) sulfanilamide: Sulfamethoxydiazine (B.A.N.)
S.2Sulfamethazine N′-(4,6-dimethyl-2- pyrimidyl)sulfanilamide: 2-(p-aminobenzenesulphonamide)-4,6-dimethylpyrimidine: sulphadimedine
S.3Sulfinpyrazone 1,2-diphenyl-4-(2-phenylsulfinilethyl)-3,5-pyrazolidinedione
S.4Sulfisoxazole 3,4-dimethyl-5-sulfanilamidoisoxazole: Sulphafurazole
T.1Tetracaine 2-dimethylaminoethyl-p-n- butylaminobenzoate: Amethocaine
T.2Thiethylperazine 2-Ethylthio-10-[3-(4- methylpiperazin-1-yl) propyl]phenothiazine
T.3Thiopropazate 2-chloro-10-[3-[1-(2-acetoxyethyl)-4-piperazinyl] propyl]phenothiazine
T.4Thioproperazine 2-Dimethylsulphamoyl-10-[3-(4-methylpiperazin-1-yl)- propyl]phenothiazine
T.5Thioridazine 10-{2-[2-(1-methylpiperidyl)] ethyl α}-2-methylthiopheno- thiazine
T.6Tranylcypromine Trans d, 1-2-phenylcyclopropyl- amine
T.7Triamterene 2,4,7-Triamino-6-phenylpteridine
T.8Triflupromazine 10-(3-dimethylaminopropyl)-2-trifluoromethylphenothiazine: Fluopremazine
T.9Trimeprazine 10-(3-dimethylamino-2-methylpropyl) phenothiazine
T.10Trimethadione 3,5,5-trimethyl-2,4-oxazolidine- dione: Troxidone
T.11Trimipramine 5-(3-Dimethylamino-2-methylpropyl)-10,11-dihydro-5H-dibenz[b,f]azepine: 5-(3′-Dimethylamino-2′-methylpropyl)iminodibenzyl
T.12Tybamate 2-Methyl-2-propyltrimethylene butylcarbamate carbamate: 2-(Hydroxymethyl)-2-methyl-pentyl butylcarbamate carbamate
V.1Vinblastine An alkaloid derived from Vinca rosea
V.2Vincristine An alkaloid derived from Vinca rosea
  • SOR/87-565, s. 1
  • SOR/88-182, s. 1
  • SOR/88-482, s. 1(F)
  • SOR/90-173, s. 1(F)
.

 No person shall sell a drug that is not labelled as required by these Regulations.

  • SOR/80-544, s. 1
.

  •  (1) The inner and outer labels of a drug shall show

    • (a) on the principal display panel

      • (i) the proper name, if any, of the drug which, if there is a brand name for the drug, shall immediately precede or follow the brand name in type not less than one-half the size of that of the brand name,

      • (ii) if there is no proper name, the common name of the drug, and

      • (iii) [Repealed, SOR/2024-238, s. 3]

      • (iv) [Repealed, SOR/2024-238, s. 3]

      • (v) in both official languages, the notation “sterile” “stérile” if the drug is required to be sterile by these Regulations;

    • (b) on the upper left quarter of the principal display panel

      • (i) the symbol “Pr” in the case of a prescription drug, but the symbol “Pr” shall not appear on the label of any other drug,

      • (ii) the symbol “Symbol consisting of a diamond shape outline in which an uppercase letter C is centred.” in a clear manner and a conspicuous colour and size, in the case of a controlled drug, other than a controlled drug contained in an agricultural implant and set out in Part III of the schedule to Part G,
      • (iii) the symbol “N” in a colour contrasting with the rest of the label or in type not less than half the size of any letters used thereon, in the case of a narcotic as defined in the Narcotic Control Regulations, and

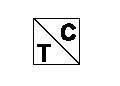
      • (iv) in the case of a targeted substance as defined in section 1 of the Benzodiazepines and Other Targeted Substances Regulations, the following symbol in a colour contrasting with the rest of the label and in type not less than half the size of any other letter used on the main panel, namely,

        Symbol consisting of a square outline divided in half from top left corner to bottom right corner. The top right half has an uppercase letter C inside and lower left half has an uppercase letter T inside.
    • (c) on any panel

      • (i) the name and address of the manufacturer of the drug,

      • (ii) the lot number of the drug,

      • (iii) adequate directions for use of the drug, except in the case of a drug to which section C.01.004.02 applies,

      • (iv) a quantitative list of the medicinal ingredients of the drug by their proper names or, if they have no proper names, by their common names, except in the case of a drug to which section C.01.004.02 applies,

      • (v) the expiration date of the drug, and

      • (vi) in the case of a new drug for extraordinary use in respect of which a notice of compliance has been issued under section C.08.004.01, the following statement, displayed in capital letters and in a legible manner:

        “HEALTH CANADA HAS AUTHORIZED THE SALE OF THIS EXTRAORDINARY USE NEW DRUG FOR [naming purpose] BASED ON LIMITED CLINICAL TESTING IN HUMANS.

        SANTÉ CANADA A AUTORISÉ LA VENTE DE CETTE DROGUE NOUVELLE POUR USAGE EXCEPTIONNEL AUX FINS DE [indication de la fin] EN SE FONDANT SUR DES ESSAIS CLINIQUES RESTREINTS CHEZ L’ÊTRE HUMAIN.”.

  • (1.1) to (1.5) [Repealed, SOR/2014-158, s. 3]

  • (2) In addition to the requirements of subsection (1), the outer label of a drug shall display the following information:

    • (a) the net amount of the drug in the container in terms of weight, measure or number;

    • (b) in the case of a drug intended for parenteral use, a quantitative list of any preservatives present therein by their proper names or, if they have no proper names, by their common names; and

    • (c) in the case of a drug for human use that contains mercury or a salt or derivative thereof as a preservative, a quantitative list of all mercurial preservatives present therein by their proper names or, if they have no proper names, by their common names.

  • (3) Where the container of a drug is too small to accommodate an inner label that conforms to the requirements of these Regulations, the inner label requirements of these Regulations do not apply to the drug in that container if

    • (a) there is an outer label that complies with the labelling requirements of these Regulations; and

    • (b) the inner label shows

      • (i) the proper name of the drug, the common name of the drug if there is no proper name or, in the case of a drug with more than one medicinal ingredient, the brand name of the drug,

      • (ii) the potency of the drug except where, in the case of a drug with more than one medicinal ingredient, the name used pursuant to subparagraph (i) for that drug is unique for a particular potency of the drug,

      • (iii) the net contents of the drug if it is not in a discrete dosage form,

      • (iv) the route of administration of the drug if other than oral,

      • (v) the lot number of the drug,

      • (vi) the name of the manufacturer of the drug,

      • (vii) the expiration date of the drug, and

      • (viii) the identification of special characteristics of the dosage form if they are not evident from the name of the drug under subparagraphs (i) or (ii).

  • (4) [Repealed, SOR/92-654, s. 2]

  • (5) Subsections (1) to (3) do not apply to a drug that is

    • (a) sold to a drug manufacturer;

    • (b) sold by a pharmacist under a prescription or by a practitioner, if its label shows suitable directions for use and meets the requirements set out in section C.01.005; or

    • (c) listed in Schedule D to the Act.

  • SOR/80-543, s. 2
  • SOR/81-334, s. 1(E)
  • SOR/85-715, s. 2
  • SOR/89-229, s. 1
  • SOR/90-216, s. 1
  • SOR/90-586, s. 1
  • SOR/92-654, s. 2
  • SOR/93-202, s. 2
  • SOR/97-228, s. 1
  • SOR/97-515, s. 1
  • SOR/2000-219, s. 1
  • SOR/2001-181, s. 4
  • SOR/2010-105, s. 2
  • SOR/2011-88, s. 1
  • SOR/2013-122, ss. 4, 5
  • SOR/2014-158, s. 3
  • SOR/2019-170, s. 18
  • SOR/2023-247, s. 2
  • SOR/2024-238, s. 3
.
  •  (1) Every label of a drug for human use in dosage form shall display the following:

    • (a) a telephone number, email address, website address, postal address or any other information that enables communication with a contact person in Canada; and

    • (b) a statement to the effect that any injury to a person’s health that is suspected of being associated with the use of the drug may be reported to the contact person.

  • (2) Subsection (1) does not apply to

    • (a) the labels of a drug that is listed in Schedule C or D to the Act and that is in dosage form; or

    • (b) the inner and outer labels of a drug to which section C.01.004.02 applies.

  • SOR/2014-158, s. 4
.

  •  (1) In addition to the requirements of section C.01.004, the outer label of a drug for human use in dosage form shall display, either one bilingual table, placed on any panel, that contains only the following information in both English and French or one table in English and one table in French, each of which is placed on any panel, that contains only the following information:

    • (a) adequate directions for use of the drug;

    • (b) a quantitative list of the drug’s medicinal ingredients by their proper names or, if they have no proper names, by their common names;

    • (c) the drug’s non-medicinal ingredients listed in alphabetical order or in descending order of predominance by their proportion in the drug, preceded by words that clearly distinguish them from the medicinal ingredients; and

    • (d) the information referred to in subsection C.01.004.01(1).

  • (2) If a package is too small to accommodate an outer label that displays one bilingual table that lists all of the drug’s non-medicinal ingredients or two unilingual tables, each of which lists all of the drug’s non-medicinal ingredients, the list of non-medicinal ingredients shall be displayed in both English and French on a tag, tape or card that is attached to the package.

  • (3) If pharmaceutical ink, a fragrance or a flavour has been added to the drug, the following expressions may be included in the list of non-medicinal ingredients to indicate that those ingredients have been added to the drug, instead of listing them individually:

    • (a) in the case where a bilingual table referred to in subsection (1) is displayed, the expressions “flavour/saveur”, “fragrance/parfum” and “pharmaceutical ink/encre pharmaceutique”; or

    • (b) in the case where two unilingual tables referred to in subsection (1) are displayed, the expressions

      • (i) “encre pharmaceutique”, “parfum” and “saveur” in the table in French, and

      • (ii) “flavour”, “fragrance” and “pharmaceutical ink” in the table in English.

  • (4) If the composition of the drug varies from one lot to another with respect to its non-medicinal ingredients,

    • (a) in the case where a bilingual table referred to in subsection (1) is displayed, the table shall include a reference to all non-medicinal ingredient alternatives that may be present in the drug, preceded by the symbol “+/–” or “±” or the expression “or/ou” or “may contain/peut contenir”; or

    • (b) in the case where two unilingual tables referred to in subsection (1) are displayed,

      • (i) the table in French shall list all non-medicinal ingredient alternatives that may be present in the drug, preceded by the symbol “+/–” or “±” or the expression “ou” or “peut contenir”, and

      • (ii) the table in English shall list all non-medicinal ingredient alternatives that may be present in the drug, preceded by the symbol “+/–” or “±” or the expression “or” or “may contain”.

  • (5) For the purposes of paragraphs (3)(a) and (4)(a), the French terms in the expressions may appear first.

  • (6) Subsections (1) to (5) do not apply to

    • (a) prescription drugs;

    • (b) drugs that are permitted to be sold without a prescription but that are to be administered only under the supervision of a practitioner; and

    • (c) drugs that are represented as being solely for use as a disinfectant on hard non-porous surfaces.

  • SOR/2014-158, s. 5
  • SOR/2017-18, s. 23
  • SOR/2018-69, s. 13
  • SOR/2021-46, s. 5
.

 In addition to the requirements of section C.01.004, the inner label of a drug to which section C.01.004.02 applies shall display on any panel

  • (a) adequate directions for use of the drug;

  • (b) a quantitative list of the drug’s medicinal ingredients by their proper names or, if they have no proper names, by their common names; and

  • (c) the information referred to in subsection C.01.004.01(1).

  • SOR/2014-158, s. 5
.
  •  (1) No person shall import a drug in dosage form into Canada for the purpose of sale unless they have in Canada a person who is responsible for the sale of the drug.

  • (2) No person who imports a drug in dosage form into Canada shall sell any lot or batch of the drug unless the name of the person who imports it, and the address of the principal place of business in Canada of the person responsible for its sale, appears on the inner and outer labels of the drug.

  • SOR/82-524, s. 1
  • SOR/93-475, s. 1
  • SOR/97-12, s. 2
.
  •  (1) The principal display panel of both the inner label and outer label of a drug in dosage form shall show the drug identification number assigned for that drug, preceded by the expression “Drug Identification Number” or “Drogue : identification numérique”, or both, or the abbreviation “DIN”.

  • (2) Subsection (1) does not apply to

    • (a) a drug in dosage form that is compounded by a pharmacist under a prescription or by a practitioner; or

    • (b) a drug in dosage form that is sold under a prescription if the following information appears on the drug’s label:

      • (i) the drug’s proper name, common name or brand name,

      • (ii) the drug’s potency, and

      • (iii) the name of the drug’s manufacturer.

  • (3) In this section and in sections C.01.005.1 and C.01.014, drug in dosage form means a drug in a form in which it is ready for use by the consumer without requiring any further manufacturing.

  • (4) and (5) [Repealed, SOR/81-248, s. 1]

  • SOR/81-248, s. 1
  • SOR/93-202, s. 3
  • SOR/98-423, s. 2
  • SOR/2001-181, s. 4
  • SOR/2017-259, s. 2
  • SOR/2018-69, s. 27
  • SOR/2018-77, s. 1
.
  •  (1) No pharmacist or practitioner shall sell a Class A opioid — including one that is compounded by a pharmacist under a prescription or by a practitioner — unless

    • (a) the drug’s package has applied to it a warning sticker that meets the specifications set out in the source document; and

    • (b) the drug is accompanied by a patient information handout that meets the specifications set out in the source document.

  • (2) Subsection (1) does not apply in respect of the sale of a Class A opioid by a pharmacist or practitioner if

    • (a) the opioid will be, or is, administered under the supervision of a practitioner; or

    • (b) the sale is to a pharmacist or practitioner.

  • (3) The following definitions apply in this section.

    Class A opioid

    Class A opioid means a drug in dosage form set out in Part A of the List of Opioids, published by the Government of Canada on its website, as amended from time to time. (opioïde de catégorie A)

    source document

    source document means the document entitled Information for Patients Concerning Opioids, published by the Government of Canada on its website, as amended from time to time. (document source)

  • SOR/2018-77, s. 2
.

 Where a package of a drug has only one label, that label shall contain all the information required by these Regulations to be shown on both the inner and the outer labels.

 No reference, direct or indirect, to the Act or to these Regulations shall be made upon any label of or in any advertisement for a drug unless such reference is a specific requirement of the Act or these Regulations.

 [Repealed, SOR/80-544, s. 2]

 If any Act of Parliament or any of its regulations prescribes a standard or grade for a drug and that standard or grade is given a name or designation in the Act or regulation, no person shall, on a label of or in any advertisement for that drug, use that name or designation unless the drug conforms with the standard or grade.

  • SOR/2017-18, s. 13
.

Amended on June 4, 2026

 If it is necessary to provide adequate directions for the safe use of a parenteral drug or prescription drug that is used in the treatment or prevention of any disease, disorder or abnormal physical state mentioned in Schedule A.1 to the Act, the disease, disorder or abnormal physical state may be mentioned on the drug’s labels, including any package insert and any document that is provided on request and that sets out supplementary information on the use of the drug, and, in that respect, the drug is exempt from subsections 3(1) and (2) of the Act.

  • SOR/2013-122, s. 6
  • SOR/2014-158, s. 6
  • SOR/2017-18, s. 21(F)
  • SOR/2018-69, s. 14(F)
  • SOR/2021-46, s. 10
  • SOR/2026-113, s. 5(F)
.
  •  (1) A drug referred to in subsection 10(2) of the Act shall be exempt from the standard for any drug contained in any publication mentioned in Schedule B to the Act to the extent that such drug differs from that standard with respect to colour, flavour, shape and size, if such difference does not interfere with any method of assay prescribed in any such publication.

  • (2) [Repealed, SOR/93-243, s. 2]

  • (3) Where a manufacturer’s standard is used for a drug, the manufacturer shall make available to the Minister, on request, details of that standard and of a method of analysis for the drug acceptable to the Minister.

  • (4) No person shall use a manufacturer’s standard for a drug that provides

    • (a) a lesser degree of purity than the highest degree of purity, or

    • (b) a greater variation in potency than the least variation in potency,

    provided for that drug in any publication mentioned in Schedule B to the Act.

  • (5) Subsection (4) does not apply in respect of new drugs, other than those listed in Schedule C to the Act.

  • SOR/93-243, s. 2
  • SOR/2018-69, ss. 31(E), 32(F)
  • SOR/2024-238, s. 4
.

 A manufacturer who makes representations on a label of a drug in oral dosage form, or in any advertisement, with respect to the site, rate or extent of release to the body of a medicinal ingredient of the drug, or the availability to the body of a medicinal ingredient of the drug, shall

  • (a) before making the representations, conduct such investigations, using an acceptable method, as may be necessary to demonstrate that the site, rate or extent of release to the body of the medicinal ingredient of the drug and the availability to the body of the medicinal ingredient of the drug, correspond to the representations; and

  • (b) on request submit the record of such investigations to the Minister.

  • SOR/89-455, s. 2
  • SOR/94-36, s. 1
  • SOR/2018-69, s. 27
.
  •  (1) Where the manufacturer of a drug is requested in writing by the Minister to submit on or before a specified day evidence with respect to a drug, the manufacturer shall make no further sales of that drug after that day unless he has submitted the evidence requested.

  • (2) If the Minister determines that the evidence submitted by a manufacturer under subsection (1) is not sufficient, he or she shall so notify the manufacturer in writing.

  • (3) Where, pursuant to subsection (2), a manufacturer is notified that the evidence with respect to a drug is not sufficient, he shall make no further sales of that drug unless he submits further evidence and is notified in writing by the Minister that that further evidence is sufficient.

  • (4) A reference in this section to evidence with respect to a drug means evidence to establish the safety of the drug under the conditions of use recommended and the effectiveness of the drug for the purposes recommended.

  • SOR/2018-69, ss. 15, 27
.

 Section C.01.013 does not apply in respect of a veterinary health product.

  • SOR/2017-76, s. 2
.

Assignment and Cancellation of Drug Identification Numbers

  •  (1) No manufacturer shall sell a drug in dosage form unless a drug identification number has been assigned for that drug and the assignment of the number has not been cancelled under section C.01.014.6.

  • (2) Subsection (1) does not apply in respect of a veterinary health product, a study drug as defined in section C.03.301 or a medicated feed as defined in subsection 1(1) of the Feeds Regulations, 2024.

  • SOR/81-248, s. 2
  • SOR/97-12, s. 3
  • SOR/2013-179, s. 1
  • SOR/2017-259, s. 3
  • SOR/2018-77, s. 3
  • SOR/2024-132, s. 85
.

  •  (1) A manufacturer of a drug may make an application for a drug identification number for that drug.

  • (2) An application under subsection (1) shall be made to the Minister in writing and shall include the following information and material:

    • (a) the name of the manufacturer of the drug as it will appear on the label;

    • (b) the pharmaceutical form in which the drug is to be sold;

    • (c) in the case of any drug other than a drug described in paragraph (d), the recommended route of administration;

    • (d) in the case of a drug for disinfection in premises, the types of premises for which its use is recommended;

    • (e) a quantitative list of the medicinal ingredients contained in the drug by their proper names or, if they have no proper names, by their common names;

    • (f) the brand name under which the drug is to be sold;

    • (g) an indication of whether the drug is for human use or veterinary use;

    • (h) the name and quantity of each colouring ingredient that is not a medicinal ingredient;

    • (i) the use or purpose for which the drug is recommended;

    • (j) the recommended dosage of the drug;

    • (k) the address of the manufacturer referred to in paragraph (a) and, where the address is outside the country, the name and address of the importer of the drug;

    • (l) the name and address of any individual, firm, partnership or corporation, other than the names and addresses referred to in paragraphs (a) and (k), that will appear on the label of the drug;

    • (m) in the case of a drug for veterinary use, the written text of every label to be used in connection with the drug, including any package insert and any document that is provided on request and that sets out supplementary information on the use of the drug;

    • (m.1) in the case of a drug for human use, mock-ups of every label to be used in connection with the drug — including any package insert and any document that is provided on request and that sets out supplementary information on the use of the drug — and mock-ups of the drug’s packages;

    • (n) the name and title of the person who signed the application and the date of signature; and

    • (o) in the case of a drug for human use, an assessment as to whether there is a likelihood that the drug will be mistaken for another drug for which a drug identification number has been assigned due to a resemblance between the brand name that is proposed to be used in respect of the drug and the brand name, common name or proper name of the other drug.

  • (3) In the case of a new drug, a new drug submission, an extraordinary use new drug submission, an abbreviated new drug submission or an abbreviated extraordinary use new drug submission filed under section C.08.002, C.08.002.01 or C.08.002.1 shall be regarded as an application for a drug identification number.

  • SOR/81-248, s. 2
  • SOR/93-202, s. 4
  • SOR/98-423, s. 3
  • SOR/2011-88, s. 2
  • SOR/2014-158, s. 7
  • SOR/2017-259, s. 4
  • SOR/2018-69, ss. 27, 33(F)
.

  •  (1) Subject to subsection (2), if a manufacturer has provided all the information and material described in subsection C.01.014.1(2) or section C.08.002, C.08.002.01 or C.08.002.1, as the case may be, in respect of a drug, the Minister shall issue to the manufacturer a document that

    • (a) sets out

      • (i) the drug identification number assigned for the drug, preceded by the abbreviation “DIN”, or

      • (ii) if there are two or more brand names for the drug, the drug identification numbers assigned by the Minister for the drug, each of which pertains to one of the brand names and is preceded by the abbreviation “DIN”; and

    • (b) contains the information referred to in paragraphs C.01.014.1(2)(a) to (f).

  • (2) The Minister may refuse to issue the document referred to in subsection (1) if he or she has reasonable grounds to believe that the product in respect of which an application referred to in section C.01.014.1 has been made

    • (a) is not a drug, or

    • (b) is a drug but its sale would cause injury to the health of the purchaser or consumer or would be a contravention of the Act or these Regulations.

  • (3) If the Minister refuses to issue the document under subsection (2), the manufacturer may submit additional information or material and request the Minister to reconsider his or her decision.

  • (4) On the basis of the additional information or material submitted under subsection (3), the Minister shall reconsider the grounds on which the refusal to issue the document was made.

  • SOR/81-248, s. 2
  • SOR/92-230, s. 1
  • SOR/98-423, s. 4
  • SOR/2011-88, s. 3
  • SOR/2017-259, s. 5
  • SOR/2018-69, ss. 16, 27, 37
.

  •  (1) The Minister may, at any time, impose terms and conditions on a drug identification number assigned for a Class B opioid or amend those terms and conditions.

  • (1.1) The Minister may, at any time, impose terms and conditions on a drug identification number assigned for a public health emergency drug or amend such terms and conditions if

    • (a) a notice of compliance was issued under section C.08.004 in respect of

      • (i) a new drug submission that was filed for the public health emergency drug under section C.08.002 and contains the statement referred to in paragraph C.08.002(2.1)(a), or

      • (ii) a supplement to any new drug submission that was filed for the public health emergency drug under section C.08.003 and contains the statement referred to in paragraph C.08.003(5)(a); or

    • (b) a notice of compliance was issued under section C.08.004 in respect of one of the following submissions or supplements that was filed in respect of the public health emergency drug on the basis of a direct or indirect comparison to another public health emergency drug referred to in paragraph (a):

      • (i) a new drug submission under section C.08.002,

      • (ii) an abbreviated new drug submission under section C.08.002.1, or

      • (iii) a supplement to a new drug submission or abbreviated new drug submission under section C.08.003.

  • (2) The Minister shall notify, in writing, the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number of any terms and conditions imposed on the drug identification number and of any amendment to those terms and conditions.

  • (3) The following definitions apply in this section.

    Class B opioid

    Class B opioid means a drug set out in Part B of the List of Opioids, published by the Government of Canada on its website, as amended from time to time. (opioïde de catégorie B)

    designated COVID-19 drug

    designated COVID-19 drug[Repealed, SOR/2024-238, s. 7]

    public health emergency drug

    public health emergency drug has the same meaning as in section C.08.001.1. (drogue pour urgence de santé publique)

  • SOR/2018-77, s. 4
  • SOR/2021-45, s. 2
  • SOR/2024-238, s. 7
.

 The manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for a drug shall, within 30 days after the day on which the drug is first sold following the issuance by the Minister of the document, date and sign the document and return it to the Minister with a statement set out on it that the information it contains is correct and with an indication of the date of that first sale.

  • SOR/81-248, s. 2
  • SOR/98-423, s. 5
  • SOR/2014-158, s. 8
  • SOR/2017-259, s. 6
  • SOR/2018-69, s. 38
.

 If the information referred to in subsection C.01.014.1(2) in respect of a drug is no longer correct owing to a change in the subject matter of the information,

  • (a) in the case of a change in the subject matter of any of the information referred to in paragraphs C.01.014.1(2)(a) to (f)

    • (i) that occurs prior to the sale of the drug, a new application shall be made, or

    • (ii) that occurs after the sale of the drug, no further sale of the drug shall be made until a new application for a drug identification number in respect of that drug is made and a number is assigned; and

  • (b) in the case of a change in the subject matter of any of the information referred to in paragraphs C.01.014.1(2)(g) to (k)

    • (i) that occurs prior to the sale of the drug, the particulars of the change shall be submitted with the return of the document referred to in section C.01.014.3, or

    • (ii) that occurs after the sale of the drug, the person to whom the drug identification number in respect of that drug was issued shall, within 30 days of the change, inform the Minister of the change.

  • SOR/81-248, s. 2
  • SOR/92-230, s. 2
  • SOR/98-423, s. 6
  • SOR/2016-139, s. 2(F)
  • SOR/2018-69, s. 27
.
  •  (1) The manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for a drug shall, annually before the first day of October and in a form established by the Minister, provide the Minister with a notification that is signed by them and that

    • (a) indicates whether any of the following circumstances apply in respect of the drug:

      • (i) as of the day on which the notification is sent,

        • (A) the manufacturer sells the drug in Canada, or

        • (B) the manufacturer has discontinued the sale of the drug in Canada, or

      • (ii) the manufacturer has not sold the drug in Canada for a period that is greater than 12 months and a portion of that period is covered by the notification; and

    • (b) subject to subsection (2), confirms that the information that the manufacturer previously submitted with respect to the drug under subsection C.01.014.1(2), paragraph C.01.014.4(b) or section C.08.002, C.08.002.01, C.08.002.1 or C.08.003, as the case may be, is correct as of the day on which the notification is sent.

  • (2) If any of the information that the manufacturer submitted under a provision referred to in paragraph (1)(b) is not correct as of the day on which the notification is sent, the manufacturer shall update that information in the notification.

  • SOR/81-248, s. 2
  • SOR/2017-259, s. 7
  • SOR/2018-69, ss. 27, 39
.
  •  (1) The Minister shall cancel the assignment of a drug identification number for a drug if

    • (a) the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number advises under section C.01.014.7 that they discontinued the sale of the drug; or

    • (c) the Minister determines that the product for which the drug identification number has been assigned is not a drug.

  • (2) The Minister may cancel the assignment of a drug identification number for a drug if

    • (a) the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number contravenes section C.01.014.5;

    • (b) the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number has been notified under section C.01.013 that the evidence that they submitted with respect to the drug is not sufficient; or

    • (c) the drug is a new drug in respect of which the notice of compliance has been suspended under section C.08.006.

  • (3) The Minister may cancel the assignment of a drug identification number for a drug if, after he or she has, under section 21.31 of the Act, ordered the holder of a therapeutic product authorization referred to in subparagraph C.01.052(1)(a)(i) or (iii) to conduct an assessment of the drug in order to provide evidence establishing that the benefits associated with the drug outweigh the risks of injury to health,

    • (a) the holder fails to comply with the order; or

    • (b) the holder complies with the order but the Minister determines that the results of the assessment are not sufficient to establish that the benefits associated with the drug outweigh the risks of injury to health.

  • (4) For greater certainty, the Minister’s power to cancel the assignment of a drug identification number

    • (a) under paragraph (2)(b) is not affected by his or her power to cancel the assignment of such a number under subsection (3); and

    • (b) under subsection (3) is not affected by his or her power to cancel the assignment of such a number under paragraph (2)(b).

  • SOR/81-248, s. 2
  • SOR/2016-139, s. 3
  • SOR/2017-259, s. 7
  • SOR/2018-69, ss. 27, 39
  • SOR/2018-84, ss. 1, 13
  • SOR/2020-262, s. 1
.

Shortages of Drugs and Interruption and Discontinuation of Sale of Drugs

Amended on June 17, 2026

  •  (1) The manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for a drug shall, within 30 days after the day on which they discontinue the sale of the drug, submit the following information to the Minister:

    • (a) the drug identification number assigned for the drug;

    • (b) the date on which the manufacturer discontinued the sale of the drug; and

    • (c) the latest expiration date of the drug that the manufacturer sold and the lot number of that drug.

  • (2) For greater certainty, subsection (1) does not remove the requirement for the manufacturer to post information under subsection C.01.014.91(1).

  • SOR/81-248, s. 2
  • SOR/2016-139, s. 5
  • SOR/2017-259, s. 8
  • SOR/2026-96, s. 2
.

Amended on June 17, 2026

 If 12 months have elapsed since the day on which the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for a drug — within the meaning of paragraph (a) of the definition drug in subsection C.01.014.8(1) — last sold the drug, the manufacturer shall so notify the Minister, in writing, within 30 days after the day on which that period ends.

  • SOR/2017-259, s. 8
  • SOR/2026-96, s. 3
.

 If the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for a drug resumes the sale of the drug after a period of 12 months has elapsed since the day on which they last sold the drug, the manufacturer shall so notify the Minister in writing within 30 days after the day on which they resume the sale of the drug.

  • SOR/2017-259, s. 8
.

Amended on June 17, 2026

  •  (1) The Minister may add a drug to Part 1 of the List of Drugs for the Purposes of Paragraphs (b) and (c) of the Definition drug in Subsection C.01.014.8(1) only if the Minister has reasonable grounds to believe that a shortage or the discontinuation of sale of the drug, if it were to occur, could present a risk of injury to human health.

  • (2) The Minister may add a class of drugs to Part 2 of the List of Drugs for the Purposes of Paragraphs (b) and (c) of the Definition drug in Subsection C.01.014.8(1) only if the Minister has reasonable grounds to believe that a shortage or the discontinuation of sale of any drug that belongs to that class, if it were to occur, could present a risk of injury to human health.

    SOR/2026-96, s. 4
.

Amended on June 17, 2026

  •  (1) In sections C.01.014.9 to C.01.014.95, other than in section C.01.014.93, drug means any of the following drugs for which a drug identification number has been assigned and not been cancelled:

    • (a) the following drugs for human use:

      • (i) drugs included in Schedule I, II, III, IV or V to the Controlled Drugs and Substances Act,

      • (ii) prescription drugs,

      • (iii) drugs that are listed in Schedule C or D to the Act, and

      • (iv) drugs that are permitted to be sold without a prescription but that are to be administered only under the supervision of a practitioner;

    • (b) drugs that are set out in Part 1 of the List of Drugs for the Purposes of Paragraphs (b) and (c) of the Definition drug in Subsection C.01.014.8(1); and

    • (c) drugs that belong to a class of drugs that is set out in Part 2 of that list.

  • (2) In sections C.01.014.9 and C.01.014.91, designated website means

    • (a) if a hyperlink is on the Government of Canada website in accordance with section C.01.014.92, the website that is maintained under contract with His Majesty in right of Canada; and

    • (b) if not, the part of the Government of Canada website that is represented as being for the purpose of posting information under sections C.01.014.9 and C.01.014.91.

  • SOR/2016-139, s. 5
  • SOR/2017-259, s. 8
  • SOR/2018-69, ss. 17, 40
  • SOR/2021-199, s. 1
  • SOR/2026-96, s. 4
.

Amended on June 17, 2026

  •  (1) Subject to subsection (5), if a shortage of a drug exists or is likely to occur, the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for the drug shall post the following information in English and French on the designated website:

    • (a) the manufacturer’s name and contact information;

    • (b) the drug identification number assigned for the drug;

    • (c) the drug’s brand name and proper name or, if it does not have a proper name, its common name;

    • (d) the proper names of the drug’s medicinal ingredients or, if they do not have proper names, their common names;

    • (e) the drug’s therapeutic classification according to the Anatomical Therapeutic Chemical classification system (ATC), established by the World Health Organization Collaborating Centre for Drug Statistics Methodology — namely the level 3 description of, and level 4 code for, the drug;

    • (f) the drug’s strength;

    • (g) the drug’s dosage form;

    • (h) the quantity of the drug contained in its package;

    • (i) the drug’s route of administration;

    • (j) the date when the shortage began or is anticipated to begin;

    • (k) the anticipated date when the manufacturer will be able to meet the demand for the drug, if they can anticipate that date; and

    • (l) the actual or anticipated reason for the shortage.

  • (2) The manufacturer shall post the information

    • (a) if they anticipate that a shortage will begin in more than six months, at least six months before the day on which they anticipate it to begin;

    • (b) if they anticipate that a shortage will begin in six months or less, within five days after the day on which they make that determination; or

    • (c) if they did not anticipate the shortage, within five days after the day on which they become aware of it.

  • (3) If any of the information that was posted by the manufacturer changes, they shall update that information on the designated website within two days after the day on which they make or become aware of the change.

  • (4) Within two days after the day on which the manufacturer is able to meet the demand for the drug, they shall post information on the designated website to that effect.

  • (5) This section does not apply in respect of a shortage of a drug that results from a decision by the manufacturer to discontinue its sale.

  • SOR/2016-139, s. 5
  • SOR/2017-259, s. 9
  • SOR/2021-199, s. 2
  • SOR/2026-96, s. 5
.

Amended on June 17, 2026

  •  (1) If the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for a drug decides to discontinue the sale of the drug, they shall post the following information in English and French on the designated website:

    • (a) the manufacturer’s name and contact information;

    • (b) the drug identification number assigned for the drug;

    • (c) the drug’s brand name and proper name or, if it does not have a proper name, its common name;

    • (d) the proper names of the drug’s medicinal ingredients or, if they do not have proper names, their common names;

    • (e) the drug’s therapeutic classification according to the Anatomical Therapeutic Chemical classification system (ATC), established by the World Health Organization Collaborating Centre for Drug Statistics Methodology — namely the level 3 description of, and level 4 code for, the drug;

    • (f) the drug’s strength;

    • (g) the drug’s dosage form;

    • (h) the quantity of the drug contained in its package;

    • (i) the drug’s route of administration;

    • (j) the date on which the manufacturer will discontinue the sale of the drug; and

    • (k) the reason for the discontinuation of sale.

  • (2) The manufacturer shall post the information

    • (a) if they decide to discontinue the sale of the drug in more than 12 months, at least 12 months before the day on which they will discontinue its sale; and

    • (b) if they decide to discontinue the sale of the drug in 12 months or less, within five days after the day on which they make that decision.

  • (3) If any of the information that was posted by the manufacturer changes, they shall update that information on the designated website within five days after the day on which they make or become aware of the change.

  • (4) For greater certainty, subsection (1) does not remove the requirement for the manufacturer to inform the Minister under section C.01.014.7.

  • SOR/2016-139, s. 5
  • SOR/2017-259, s. 10
  • SOR/2026-96, s. 6
.

Amended on June 17, 2026

 If a person maintains a website under contract with His Majesty in right of Canada for the purpose of posting information under sections C.01.014.9 and C.01.014.91, the Minister shall ensure that a hyperlink to that website is on the Government of Canada website.

  • SOR/2016-139, s. 5
  • SOR/2017-259, s. 11
  • SOR/2026-96, s. 7
.

Amended on June 17, 2026

  •  (1) The Minister may request that the manufacturer to whom a document was issued under subsection C.01.014.2(1) that sets out the drug identification number assigned for a drug — or any person who holds an establishment licence in respect of a drug for which a drug identification number has been assigned and not been cancelled — provide the Minister with information that is in their control if the Minister has reasonable grounds to believe that

    • (a) there is a shortage or risk of shortage of the drug;

    • (a.1) a shortage of the drug, if it were to occur, could present a risk of injury to human health;

    • (b) the information is necessary to establish or assess

      • (i) the existence of a shortage or risk of shortage of the drug,

      • (ii) the reason for a shortage or risk of shortage of the drug,

      • (iii) the effects or potential effects on human health of a shortage of the drug,

      • (iv) measures that could be taken to prevent or mitigate shortage risks associated with the drug, or

      • (v) measures that could be taken to mitigate the impacts of a shortage of the drug; and

    • (c) the manufacturer or licensee will not provide the information without a legal obligation to do so.

  • (2) The manufacturer or licensee shall provide the requested information in the time, form and manner specified by the Minister.

  • SOR/2021-199, s. 3
  • SOR/2026-96, s. 8
.

Amended on June 17, 2026

 No person who holds an establishment licence shall distribute a drug for consumption or use outside Canada unless the licensee has reasonable grounds to believe that the distribution will not cause or exacerbate a shortage of the drug.

  • SOR/2021-199, s. 3
  • SOR/2026-96, s. 9
.

Amended on June 17, 2026

  •  (1) If a person who holds an establishment licence distributes a drug for consumption or use outside Canada, the licensee shall immediately create a detailed record of the information that they relied on to determine that the distribution of the drug is not prohibited by section C.01.014.94.

  • (2) The licensee shall retain the record for at least one year after the latest expiration date of the drug that they distributed.

  • SOR/2021-199, s. 3
  • SOR/2026-96, s. 10
.

Tablet Disintegration Times

  •  (1) Subject to subsection (2), no person shall sell for human use a drug in the form of a tablet that is intended to be swallowed whole unless, when tested by the official method DO-25, Determination of the Disintegration Time of Tablets, dated July 5, 1989,

    • (a) in the case of an uncoated tablet, the tablet disintegrates in not more than 45 minutes;

    • (b) in the case of a plain coated tablet, the tablet disintegrates in not more than 60 minutes; and

    • (c) in the case where the label of the drug indicates that the tablet carries an enteric coating or a coating designed to serve a purpose similar to that of an enteric coating, the tablet does not disintegrate when exposed for 60 minutes to simulated gastric fluid, but when it is subsequently exposed for a continuous period to simulated intestinal fluid, the tablet disintegrates in not more than 60 minutes.

  • (2) Subsection (1) does not apply in respect of a drug in the form of a tablet where

    • (a) a notice of compliance in respect of the drug in the form of a tablet has been issued under section C.08.004 or C.08.004.01;

    • (b) [Repealed, SOR/98-423, s. 7]

    • (c) a dissolution or disintegration test for the drug in the form of a tablet is prescribed in Division 6 of this Part;

    • (d) the drug is labelled as complying with a standard contained in a publication referred to in Schedule B to the Act;

    • (e) the drug has been demonstrated by an acceptable method to be available to the body; or

    • (f) representations regarding the drug are made on its label, or in any advertisement, with respect to the site, rate or extent of release to the body of a medicinal ingredient of that drug, or the availability to the body of a medicinal ingredient of that drug.

  • SOR/89-429, s. 2
  • SOR/89-455, s. 3
  • SOR/94-36, s. 2
  • SOR/98-423, s. 7
  • SOR/2011-88, s. 4
.

Prohibition

 No manufacturer shall sell a drug unless the manufacturer complies with the conditions set out in sections C.01.017 to C.01.019.

  • SOR/95-521, s. 2
  • SOR/2011-31, s. 1
.

Serious Adverse Drug Reaction Reporting — Manufacturers

 The manufacturer shall submit to the Minister a report of all information relating to the following serious adverse drug reactions within 15 days after receiving or becoming aware of the information, whichever occurs first:

  • (a) any serious adverse drug reaction that has occurred in Canada with respect to the drug; and

  • (b) any serious unexpected adverse drug reaction that has occurred outside Canada with respect to the drug.

  • SOR/95-521, s. 2
  • SOR/2011-31, s. 1
.

Annual Summary Report and Case Reports

  •  (1) The manufacturer shall prepare an annual summary report of all information relating to adverse drug reactions and serious adverse drug reactions to the drug that it received or became aware of during the previous 12 months.

  • (2) The annual summary report shall contain a concise, critical analysis of the adverse drug reactions and serious adverse drug reactions to the drug.

  • (3) In preparing the annual summary report, the manufacturer shall determine, on the basis of the analysis referred to in subsection (2), whether there has been a significant change in what is known about the risks and benefits of the drug during the period covered by the report and shall include its conclusions in this regard in the summary report.

  • (4) If, in preparing the annual summary report, the manufacturer concludes that there has been a significant change, it shall notify the Minister without delay, in writing, unless this has already been done.

  • (5) The Minister may, for the purposes of assessing the safety and effectiveness of the drug, request in writing that the manufacturer submit to the Minister one or both of the following:

    • (a) the annual summary reports;

    • (b) the case reports relating to the adverse drug reactions and serious adverse drug reactions to the drug that are known to the manufacturer.

  • (6) The Minister shall, after giving the manufacturer an opportunity to be heard, specify a period for the submission of the annual summary reports or case reports, or both, that is reasonable in the circumstances, and the manufacturer shall submit the reports within that period.

  • SOR/2011-31, s. 1
.

 Section C.01.018 does not apply in respect of a veterinary health product.

  • SOR/2017-76, s. 4
.

Issue-Related Summary Report

  •  (1) The Minister may, for the purposes of assessing the safety and effectiveness of the drug, request in writing that the manufacturer submit to the Minister an issue-related summary report.

  • (2) The report shall contain a concise, critical analysis of the adverse drug reactions and serious adverse drug reactions to the drug, as well as case reports of all or specified adverse drug reactions and serious adverse drug reactions to the drug that are known to the manufacturer in respect of the issue that the Minister directs the manufacturer to analyze in the report.

  • (3) The Minister shall, after giving the manufacturer an opportunity to be heard, specify a period for the submission of the report that is reasonable in the circumstances. The Minister may only specify a period that is less than 30 days if the Minister needs the information in the report to determine whether the drug poses a serious and imminent risk to human health.

  • (4) The manufacturer shall submit the report within the specified period.

  • SOR/2011-31, s. 1
  • SOR/2017-18, s. 14
.

 Section C.01.019 does not apply in respect of a veterinary health product.

  • SOR/2017-76, s. 5
.

Maintenance of Records

  •  (1) The manufacturer shall maintain records of the reports and case reports referred to in sections C.01.017 to C.01.019.

  • (2) The manufacturer shall retain the records for 25 years after the day on which they were created.

  • SOR/2011-31, s. 1
.

Provision of Information Under Section 21.8 of Act

Amended on June 17, 2026

  •  (1) For the purposes of section 21.8 of the Act, hospitals are the prescribed health care institutions that shall provide information that is in their control to the Minister about a serious adverse drug reaction.

  • (2) The following prescribed information about a serious adverse drug reaction that is in a hospital’s control shall be provided to the Minister in writing within 30 days after the day on which the serious adverse drug reaction is first documented within the hospital:

    • (a) the name of the hospital and the contact information of a representative of that hospital;

    • (b) the drug’s brand name, proper name or common name;

    • (c) in the case of a drug imported under subsection C.10.001(2) or section C.10.006 or sold under subsection C.10.007.1(1), the identifying code or number of the drug, if any, assigned in the country in which the drug was authorized for sale;

    • (c.1) in the case of a drug whose sale has been authorized under subsection C.11.003(1), its identifying name, code, number or mark;

    • (d) the drug identification number assigned for the drug, if applicable;

    • (e) the patient’s age and sex;

    • (f) a description of the serious adverse drug reaction;

    • (g) the date on which the serious adverse drug reaction was first documented;

    • (h) the date on which the patient first used the drug and, if applicable, the date on which the patient stopped using the drug;

    • (i) the date on which the serious adverse drug reaction first occurred and, if applicable, the date on which the patient’s health was restored to its state prior to the reaction;

    • (j) any medical condition of the patient that directly relates to the serious adverse drug reaction;

    • (k) any concomitant therapeutic products used by the patient; and

    • (l) the effect of the serious adverse drug reaction on the patient’s health.

  • (3) A hospital is exempt from section 21.8 of the Act in respect of the reporting of information referred to in subsection (2) if

    • (a) the hospital does not have in its control all of the information referred to in paragraphs (2)(b), (c), (e) and (f) in respect of the serious adverse drug reaction; or

    • (b) the serious adverse drug reaction relates only to any of the following drugs:

      • (i) a vaccine that was administered under a routine immunization program of a province,

      • (ii) a drug that is authorized for sale under Division 5 of this Part, or

      • (iii) a drug that was sold under subsection C.08.011(1).

  • (4) In this section, hospital means a facility

    • (a) that is licensed, approved or designated as a hospital by a province in accordance with the laws of the province to provide care or treatment to persons suffering from any form of disease or illness; or

    • (b) that is operated by the Government of Canada and that provides health services to in-patients.

  • SOR/2019-190, s. 2
  • SOR/2021-199, s. 4
  • SOR/2023-18, s. 1
  • SOR/2026-96, s. 11
.

Limits of Drug Dosage

Amended on June 4, 2026

 Except as provided in these Regulations, no person shall sell a drug for human use listed in the following table unless both the inner and the outer labels other than the inner label of a single dose container carry a statement of

  • (a) the quantitative content of the drug,

  • (b) the recommended single and daily adult dose designated as such, except for

    • (i) preparations solely for external use, or

    • (ii) preparations solely for children’s use; and

  • (c) adequate directions for use when the drug is recommended for children which shall be either

    • (i) the statement “CHILDREN: As directed by the physician”, or

    • (ii) reduced maximum single and daily doses which shall not exceed the following:

      Age in yearsProportion of adult dose
      10 - 14one-half
      5 - 9one-fourth
      2 - 4one-sixth
      under 2 yearsas directed by a physician

TABLE

Table of Limits of Drug Dosage for Adults

ItemExternal UseInternal Use
——
Maximum LimitMaximum Dosage Unless otherwise stated, doses are in milligrams
Per centSingleDaily
Acetaminophen —6504.0 g
Acetanilide and derivatives (except N-Acetyl-ρ-amino phenol) —65195
Acetylsalicylic Acid —6504.0 g
Aconitine, its preparations and derivatives 0.20.10.1
Adonis vernalis —65195
Amylocaine, its salts and derivatives when sold or recommended for opthalmic use 0.00.00.0
Amylocaine Hydrochloride, except when sold or recommended for ophthalmic use 1.00.00.0
Antimony, compounds of —3.313
Atropine, Methylatropine, and their salts 1.00.130.44
Belladonna and its preparations, on the basis of belladonna alkaloids 0.3750.130.44
Benzene (Benzol) ———
Benzocaine 8.0195585
Beta-Naphthol —195585
Butacaine, its salts and derivatives when sold or recommended for ophthalmic use 0.00.00.0
Butacaine Sulphate, except when sold or recommended for opthalmic use 1.00.00.0
Cadexomer Iodine 0.00.00.0
Cantharides, cantharidin, and their preparations, on the basis of cantharidin, except blisters 0.030.00.0
Cantharides, blisters only 0.20.00.0
Cedar Oil 25.00.00.0
Chlorbutol (not more often than every 4 hours) —325975
Choline Salicylate —8705.22 g
Cinchocaine Hydrochloride, except suppositories 1.00.00.0
Cinchocaine Hydrochloride, suppositories only —1111
Colchicine and its salts —0.551.65
Colchicum and its preparations, on the basis of colchicine —0.270.81
Croton Oil 10.00.00.0
Cyproheptadine and its salts — when sold or recommended for the promotion of weight gain—0.00.0
Ephedrine and its salts —1132.5
Ephedrine and its salts, sprays 1.0——
Epinephrine and its salts, sprays 1.0——
Gelseminine (Gelsemine) and its salts (not to be repeated within 4 hours) —0.551.65
Gelsemium and its preparations, on the basis of the crude drug —16.248.6
Hydrocyanic (Prussic) Acid as 2 per cent solution —0.062 ml0.31 ml
Hydroquinone 2.0——
Hyoscine (Scopolamine) and its salts 0.50.3250.975
Hyoscine aminoxide hydrobromide 0.50.3250.975
Hyoscyamine and its salts —0.3250.975
Hyoscyamus and its preparations, on the basis of hyoscyamus alkaloids —0.0730.22
Lobelia and its preparations, on the basis of the crude drug —130390
Lobeline and its salts —2.06.0
Magnesium Salicylate —6504.0 g
Methyl Salicylate 30——
Methylene Blue —130390
Phenacetin —6501.95 g
Phenazone and compounds thereof —325975
Phenol 2.032.5260
Phenylpropanolamine when sold or recommended as an appetite depressent —0.00.0
Phosphorus —0.00.0
Podophyllin 0.00.00.0
Potassium Chlorate —325975
Potassium Chlorate, gargle 2.5——
Procaine and its salts ———
Proxymetacaine, its salts and derivatives when sold or recommended for ophthalmic use 0.00.00.0
Salicylamide —9752.925 g
Santonin —65130
Selenium and its compounds 2.50.00.0
Sodium Chlorate —325975
Sodium Fluoride —0.10.1
Sodium Salicylate —6504.0 g
Squill and its preparations, on the basis of crude drug —32.597.5
Stramonium and its preparations, on the basis of stramonium alkaloids —0.160.65
Strychnine and its salts —0.00.0
Tannic Acid —1501 000
Tetracaine, its salts and derivatives when sold or recommended for ophthalmic use 0.00.00.0
Thiocyanates 0.00.00.0
Urethane 0.00.00.0

Where drugs having similar physiological actions occur in combination, the dosage of each shall be proportionately reduced.

Accurate dosagesmay be expressed in either metric units or imperial units. If the dosage is expressed in both systems, then an approximation may be used for one expression, but such approximation must precede or follow the accurate statement by which the product will be judged and must be in brackets.

  • SOR/78-422, s. 1
  • SOR/80-544, s. 3
  • SOR/84-145, s. 1
  • SOR/85-715, s. 3
  • SOR/85-966, s. 2
  • SOR/88-94, s. 1
  • SOR/89-229, s. 2
  • SOR/89-548, s. 1
  • SOR/2026-113, s. 6
.

 Notwithstanding paragraph C.01.021(b), the recommended single and daily dosage of a drug

  • (a) intended to be burned and the smoke inhaled may be increased to 10 times the oral dose, and

  • (b) intended for use as suppositories may be increased to 33 1/3 per cent in excess of the oral dose.

  •  (1) Sections C.01.021 and C.01.022 do not apply to

    • (a) a drug sold to a drug manufacturer; or

    • (b) a drug sold on prescription.

  • (2) Paragraph C.01.021(c) does not apply to

    • (a) acetaminophen;

    • (b) acetylsalicylic acid;

    • (c) magnesium salicylate;

    • (d) sodium salicylate; or

    • (e) choline salicylate.

  • (3) Where a drug mentioned in any of paragraphs (2)(a) to (d) is recommended for children, no person shall sell the drug for human use unless both the inner and the outer labels carry a statement that it is recommended

    • (a) that the drug be used as directed by a physician; or

    • (b) that the maximum doses of the drug not exceed the amounts set out in the following table and that single doses not be administered more frequently than every four hours.

    TABLE

    Maximum Dose

    ItemColumn IColumn IIColumn IIIColumn IVColumn VColumn VIColumn VII
    AgeMaximum Children’s Dose (80 mg units) Acetaminophen DropsMaximum Children’s Dose (80 mg units)Maximum Children’s Dose (160 mg units) AcetaminophenMaximum Adult’s Dose (325 mg units)Maximum Single Dose (mg)Maximum Daily Dose (mg)
    111 to under 12 years—631.54802 400
    29 to under 11 years—52.51.254002 000
    36 to under 9 years—4213201 600
    44 to under 6 years—31.5—2401 200
    52 to under 4 years—21—160800
    61 to under 2 years1.5 or as directed by a physician———120600
    74 months to under 1 year1 or as directed by a physician———80400
    80 to under 4 months0.5 or as directed by a physician———40200
  • (4) Where choline salicylate is recommended for children, no person shall sell the drug for human use unless both the inner and the outer labels carry a statement that it is recommended

    • (a) that the drug be used as directed by physician; or

    • (b) that the maximum doses of the drug not exceed the amounts set out in the following table and that single doses not be administered more frequently than every four hours.

    TABLE

    MAXIMUM DOSE
    Age (Years)Adult Dosage Units (435 mg)Single Dose (mg)Maximum Daily Dose (mg)
    11 to under 121 ½6603 300
    9 to under 111 ¼5502 750
    6 to under 914402 200
    4 to under 6¾3301 650
    2 to under 4½2201 100
    Under 2As directed by physician
  • SOR/84-145, s. 2
  • SOR/90-587, s. 1
.

 Both the inner and the outer labels of a drug that carry a recommended single or daily dosage or a statement of concentration in excess of the limits provided by section C.01.021 shall carry a caution that the product is to be used only on the advice of a physician.

 The provisions of section C.01.025 do not apply to

  • (a) a drug sold on prescription, or

  • (b) the inner label of a single-dose container.

  •  (1) Where a person advertises to the general public a drug for human use, the person shall not make any representation other than with respect to the brand name, proper name, common name, price and quantity of the drug if it

    • (a) contains a drug set out in the table to section C.01.021; and

    • (b) carries on its label

      • (i) a statement of the recommended single or daily adult dosage that results in a single or daily adult dosage of the drug referred to in paragraph (a) in excess of the maximum dosage set out in the table to section C.01.021 for that drug, or

      • (ii) a statement that shows a concentration of the drug referred to in paragraph (a) in excess of the maximum limit set out in the table to section C.01.021 for that drug.

  • (2) Subsection (1) does not apply to products containing

    • (a) acetaminophen;

    • (b) acetylsalicylic acid;

    • (c) choline salicylate;

    • (d) magnesium salicylate; or

    • (e) sodium salicylate.

  • (3) [Repealed, SOR/94-409, s. 1]

  • (4) Where a person advertises to the general public a drug for human use that contains acetylsalicylic acid, the person shall not make any representation with respect to its administration to or use by children or teenagers.

  • SOR/81-358, s. 1
  • SOR/84-145, s. 3
  • SOR/85-715, s. 4(F)
  • SOR/85-966, s. 3
  • SOR/93-202, s. 5
  • SOR/93-411, s. 1
  • SOR/94-409, s. 1
.

Cautionary Statements and Child Resistant Packages

  •  (1) Subject to subsection (2), the inner and outer labels of a drug that contains

    • (a) acetylsalicylic acid or any of its salts or derivatives, salicylic acid or a salt thereof, or salicylamide, where the drug is recommended for children, shall carry a cautionary statement to the effect that the drug should not be administered to a child under two years of age except on the advice of a physician;

    • (b) boric acid or sodium borate as a medicinal ingredient shall carry a cautionary statement to the effect that the drug should not be administered to a child under three years of age;

    • (c) hyoscine (scopolamine) or a salt thereof shall carry a cautionary statement to the effect that the drug should not be used by persons suffering from glaucoma or where it causes blurring of the vision or pressure pain within the eye;

    • (d) phenacetin, either singly or in combination with other drugs, shall carry the following cautionary statement:

      “CAUTION: May be injurious if taken in large doses or for a long time. Do not exceed the recommended dose without consulting a physician.”; or

    • (e) acetylsalicylic acid for internal use shall carry a cautionary statement to the effect that the drug should not be administered to or used by children or teenagers who have chicken pox or manifest flu symptoms before a physician or pharmacist is consulted about Reye’s syndrome, which statement shall also refer to the fact that Reye’s syndrome is a rare and serious illness.

  • (2) Subsection (1) does not apply to a drug that is

  • SOR/86-93, s. 2
  • SOR/88-323, s. 2(F)
  • SOR/93-411, s. 2
  • SOR/2013-122, s. 7
.
  • SOR/86-93, s. 2
  • SOR/87-484, s. 2
  • SOR/88-323, s. 3(F)
  • SOR/90-587, s. 2
  • SOR/93-468, s. 1
  • SOR/2018-132, s. 1
.

 [Repealed, SOR/2003-196, s. 104]

  •  (1) Subject to section C.01.031.2,

    • (a) no person shall sell a drug described in subsection C.01.029(1) unless

      • (i) where the drug is recommended solely for children, it is packaged in a child resistant package,

      • (ii) where the drug is not recommended solely for children, other than a drug referred to in subparagraph (iii), at least one of the sizes of packages available for sale is packaged in a child resistant package, and

      • (iii) where the drug is a vaping product referred to in paragraph C.01.029(1)(a.1), it is packaged in a child resistant package; and

    • (b) where a drug described in subsection C.01.029(1) is packaged in a package that is not a child resistant package, the outer label shall carry a statement that the drug is available in a child resistant package.

  • (2) Subsection C.01.031.2(2) and paragraph C.01.031.2(3)(a) do not apply to a drug referred to in paragraph C.01.029(1)(a.1).

  • SOR/86-93, s. 2
  • SOR/87-16, s. 1
  • SOR/93-468, s. 2
  • SOR/2018-132, s. 2
.

  •  (1) Sections C.01.029 to C.01.031 do not apply to a drug that is

    • (a) a prescription drug or a drug that is required to be sold under a prescription by Part G, the Benzodiazepines and Other Targeted Substances Regulations or the Narcotic Control Regulations;

    • (b) intended for parenteral use only;

    • (c) in effervescent or powder form;

    • (d) in suppository form;

    • (e) intended for topical use, unless it is a liquid preparation containing more than five per cent alkyl salicylates;

    • (f) packaged in a non-reclosable package containing not more than two adult standard dosage units per package; or

    • (g) in toothpaste form.

  • (2) Sections C.01.029 to C.01.031 do not apply to a drug that is repackaged by a pharmacist or practitioner at the time of sale.

  • (3) Section C.01.031 does not apply to a drug that is

    • (a) sold only in containers that have roll-on or spray applicators or permanently installed wick applicators;

    • (b) sold for exclusive use in animals other than household pets; or

    • (c) intended solely for use in dentists’ offices, or packaged for hospital use only.

  • SOR/86-93, s. 2
  • SOR/87-484, s. 4
  • SOR/88-323, s. 5(F)
  • SOR/93-468, s. 3
  • SOR/2013-122, s. 8
.

 No person shall sell a corticosteroid drug for ophthalmic use unless

  • (a) the outer label or the package insert carries, as part of the directions for use, the following statements:

    “Contraindications

    Viral diseases of the cornea and conjunctiva;

    Tuberculosis of the eye;

    Fungal diseases of the eye;

    Acute purulent untreated infections of the eye, which, like other diseases caused by micro-organisms, may be masked or enhanced by the presence of the steroid.

    Side Effects

    Extended ophthalmic use of corticosteroid drugs may cause increased intraocular pressure in certain individuals and in those diseases causing thinning of the cornea, perforation has been known to occur.”;

    and

  • (b) the inner label carries the statements required by paragraph (a) or instructions to see the outer label or package insert for information about contraindications and side effects.

  • SOR/2018-69, s. 18(F)
.

 Section C.01.032 does not apply to a corticosteroid drug that is sold by a pharmacist under a prescription.

  • SOR/2013-122, s. 9
.

 No person shall disseminate to a practitioner promotional literature about corticosteroid drugs for ophthalmic use unless the statements required by paragraph C.01.032(a) are included in that literature.

 Sections C.01.032 and C.01.034 do not apply to a drug sold solely for veterinary use.

Miscellaneous

  •  (1) No manufacturer or importer shall sell

    • (a) a drug that contains phenacetin in combination with any salt or derivative of salicylic acid;

    • (b) a drug for human use that contains

      • (i) oxyphenisatin,

      • (ii) oxyphenisatin acetate, or

      • (iii) phenisatin; or

    • (c) a drug for human use that contains mercury or a salt or derivative thereof, unless the drug is

      • (i) a drug described in Schedule C or D to the Act, or

      • (ii) one of the following drugs, namely,

        • (A) an ophthalmic drug or other drug to be used in the area of the eye,

        • (B) a drug for nasal administration,

        • (C) a drug for otic administration, or

        • (D) a drug for parenteral administration that is packaged in a multi-dose container,

      in which the mercury or the salt or derivative thereof is present as a preservative and the manufacturer or importer has submitted evidence to the Minister demonstrating that the only satisfactory way to maintain the sterility or stability of the drug is to use that preservative.

  • (2) For the purpose of clause (1)(c)(ii)(A), area of the eye means the area bounded by the supraorbital and infraorbital ridges and includes the eyebrows, the skin underlying the eyebrows, the eyelids, the eyelashes, the conjunctival sac of the eye, the eyeball and the soft tissue that lies below the eye and within the infraorbital ridge.

  • SOR/78-423, s. 2
  • SOR/86-93, s. 3
  • SOR/89-229, s. 3
  • SOR/2018-69, s. 27
.

 No person shall sell, or advertise for sale, nitrous oxide to the general public.

  • SOR/78-875, s. 1
.
  •  (1) No person shall sell to the general public a drug that is recommended solely for children if the package in which the drug is sold contains

    • (a) more than 1.92 g of salicylamide or salicylic acid or the equivalent quantity of a salt of salicylic acid;

    • (b) more than 1.92 g of acetylsalicylic acid or the equivalent quantity of a salt or derivative thereof;

    • (c) more than 3.2 g of acetaminophen in 160 mg dosage units; or

    • (d) more than 1.92 g of acetaminophen in 80 mg dosage units.

  • (2) Subsection (1) does not apply to a drug dispensed pursuant to a prescription.

  • SOR/86-93, s. 4
  • SOR/87-484, s. 5
  • SOR/88-323, s. 6
  • SOR/90-587, s. 3
.

 A drug for human use is adulterated if it contains

  • (a) Strychnine or any of its salts;

  • (b) extracts or tinctures of

    • (i) Strychnos nux vomica,

    • (ii) Strychnos Ignatii, or

    • (iii) a Strychnos species containing strychnine, other than those species mentioned in subparagraphs (i) and (ii);

  • (c) Methapyrilene or any of its salts;

  • (d) Echimidine or any of its salts; or

  • (e) any of the following plant species or extracts or tinctures thereof:

    • (i) Symphytum asperum,

    • (ii) Symphytum x uplandicum, or

    • (iii) any other plant species containing echimidine.

  • SOR/79-512, s. 1
  • SOR/88-173, s. 1
.

 In vitro diagnostic products that are or contain drugs other than drugs listed in Schedule E to the Act, and drugs listed in Schedule D to the Act that are labelled for veterinary use only, are exempt from the application of this Part.

  • SOR/97-12, s. 4
.

 No manufacturer or importer shall sell a drug for human use that contains as an ingredient

  • (a) chloroform; or

  • (b) arsenic or any of its salts or derivatives, other than arsenic trioxide.

  • SOR/89-229, s. 4
  • SOR/2013-113, s. 1
.

 No manufacturer shall use methyl salicylate as a medicinal ingredient in a drug for internal use in humans.

  • SOR/78-422, s. 2
  • SOR/78-801, s. 1
  • SOR/81-334, s. 2(F)
  • SOR/89-176, s. 1
  • SOR/92-662, s. 1
.

Colouring Agents

  •  (1) No manufacturer shall use a colouring agent in a drug other than a colouring agent listed in subsections (3) and (4).

  • (2) No person shall import for sale a drug that contains a colouring agent other than a colouring agent listed in subsections (3) and (4).

  • (2.1) The following definitions apply in subsections (3) to (4.1):

    C.I. (indication of the number)

    C.I. (indication of the number) means the designation used to identify a colouring agent in the Colour Index published by The Society of Dyers and Colourists, as amended from time to time; (C.I. (indication du numéro))

    D & C (indication of the colour and the number)

    D & C (indication of the colour and the number) means the designation used to identify, in accordance with the Code of Federal Regulations of the United States, a colouring agent that can be used in the United States in drugs and cosmetics; (D&C (indication de la couleur et du numéro))

    FD & C (indication of the colour and the number)

    FD & C (indication of the colour and the number) means the designation used to identify, in accordance with the Code of Federal Regulations of the United States, a colouring agent that can be used in the United States in food, drugs and cosmetics. (FD&C (indication de la couleur et du numéro))

  • (3) The following colouring agents are permitted in drugs for internal and external use, namely,

    • (a) ACID FUCHSIN D (D & C Red No. 33; C.I. No. 17200),

      • ALIZARIN CYANINE GREEN F (D & C Green No. 5; C.I. No. 61570),
      • ALLURA RED AC (FD & C Red No. 40; C.I. No. 16035),
      • AMARANTH (Delisted FD & C Red No. 2; C.I. No. 16185),
      • ANTHOCYANIN DERIVED FROM JUICE EXPRESSED FROM FRESH EDIBLE FRUITS OR VEGETABLES,
      • ß-APO-81-CAROTENAL (C.I. No. 40820),
      • BRILLIANT BLUE FCF SODIUM SALT (FD & C Blue No. 1; C.I. No. 42090),
      • BRILLIANT BLUE FCF AMMONIUM SALT (D & C Blue No. 4; C.I. No. 42090),
      • CANTHAXANTHIN (C.I. No. 40850),
      • CARAMEL,
      • CARBON BLACK (C.I. No. 77266),
      • CARMINE (C.I. No. 75470),
      • CARMOISINE (Delisted Ext. D & C Red No. 10; C.I. No. 14720),
      • ß-CAROTENE (C.I. No. 40800),
      • CHLOROPHYLL (C.I. No. 75810),
      • EOSIN YS ACID FORM (D & C Red No. 21; C.I. No. 45380:2),
      • EOSIN YS SODIUM SALT (D & C Red No. 22; C.I. No. 45380),
      • ERYTHROSINE (FD & C Red No. 3; C.I. No. 45430),
      • FAST GREEN FCF (FD & C Green No. 3; C.I. No. 42053),
      • FLAMING RED (D & C Red No. 36; C.I. No. 12085),
      • HELINDONE PINK CN (D & C Red No. 30; C.I. No. 73360),
      • INDIGO (D & C Blue No. 6; C.I. No. 73000),
      • INDIGOTINE (FD & C Blue No. 2; C.I. No. 73015),
      • IRON OXIDES (C.I. Nos. 77489, 77491, 77492, 77499),
      • LITHOL RUBIN B SODIUM SALT (D & C Red No. 6; C.I. No. 15850),
      • LITHOL RUBIN B CALCIUM SALT (D & C Red No. 7; C.I. No. 15850:1),
      • PHLOXINE B ACID FORM (D & C Red No. 27; C.I. No. 45410:1),
      • PHLOXINE B SODIUM SALT (D & C Red No. 28; C.I. No. 45410),
      • PONCEAU 4R (C.I. No. 16255),
      • PONCEAU SX (FD & C Red No. 4; C.I. No. 14700),
      • QUINOLINE YELLOW WS (D & C Yellow No. 10; C.I. No. 47005),
      • RIBOFLAVIN,
      • SUNSET YELLOW FCF (FD & C Yellow No. 6; C.I. No. 15985),
      • TARTRAZINE (FD & C Yellow No. 5; C.I. No. 19140),
      • TITANIUM DIOXIDE (C.I. No. 77891);
    • (b) preparations made by extending any of the colouring agents listed in paragraph (a) on a substratum of

      • (i) alumina,

      • (ii) blanc fixe,

      • (iii) gloss white,

      • (iv) clay,

      • (v) zinc oxide,

      • (vi) talc,

      • (vii) rosin,

      • (viii) aluminum benzoate,

      • (ix) calcium carbonate, or

      • (x) any combination of the substances listed in subparagraphs (i) to (ix); and

    • (c) preparations made by extending any sodium, potassium, aluminum, barium, calcium, strontium or zirconium salt of any of the colouring agents listed in paragraph (a) on a substratum of

      • (i) alumina,

      • (ii) blanc fixe,

      • (iii) gloss white,

      • (iv) clay,

      • (v) zinc oxide,

      • (vi) talc,

      • (vii) rosin,

      • (viii) aluminum benzoate,

      • (ix) calcium carbonate, or

      • (x) any combination of the substances listed in subparagraphs (i) to (ix).

  • (4) The following colouring agents are permitted in drugs for external use, namely,

    • (a) ACID VIOLET 43 (Ext. D & C Violet No. 2; C.I. No. 60730),

      • ALIZUROL PURPLE SS (D&C Violet No. 2; C.I. No. 60725),
      • ANNATTO (C.I. No. 75120),
      • BISMUTH OXYCHLORIDE (C.I. No. 77163),
      • CHROMIUM HYDROXIDE GREEN (PIGMENT GREEN 18 (C.I. No. 77289)),
      • DEEP MAROON (D&C Red No. 34; C.I. No. 15880:1),
      • DIBROMOFLUORESCEIN (SOLVENT RED 72 (C.I. No. 45370:1); ORANGE No. 5 (D & C Orange No. 5)),
      • FERRIC FERROCYANIDE (C.I. No. 77510),
      • GUANINE (C.I. No. 75170),
      • MANGANESE VIOLET (C.I. No. 77742),
      • MICA (C.I. No. 77019),
      • ORANGE II (D&C Orange No. 4; C.I. No. 15510),
      • PYRANINE CONCENTRATED (D&C Green No. 8; C.I. No. 59040),
      • QUINIZARIN GREEN SS (D&C Green No. 6; C.I. No. 61565),
      • TONEY RED (D&C Red No. 17; C.I. No. 26100),
      • URANINE ACID FORM (D&C Yellow No. 7; C.I. No. 45350:1),
      • URANINE SODIUM SALT (D&C Yellow No. 8; C.I. No. 45350);
      • ZINC OXIDE (C.I. No. 77947);
    • (b) preparations made by extending any of the colouring agents listed in paragraph (a) on a substratum of

      • (i) alumina,

      • (ii) blanc fixe,

      • (iii) gloss white,

      • (iv) clay,

      • (v) zinc oxide,

      • (vi) talc,

      • (vii) rosin,

      • (viii) aluminum benzoate,

      • (ix) calcium carbonate, or

      • (x) any combination of the substances listed in subparagraphs (i) to (ix); and

    • (c) preparations made by extending any sodium, potassium, aluminum, barium, calcium, strontium or zirconium salt of any of the colouring agents listed in paragraph (a) on a substratum of

      • (i) alumina,

      • (ii) blanc fixe,

      • (iii) gloss white,

      • (iv) clay,

      • (v) zinc oxide,

      • (vi) talc,

      • (vii) rosin,

      • (viii) aluminum benzoate,

      • (ix) calcium carbonate, or

      • (x) any combination of the substances listed in subparagraphs (i) to (ix).

  • (4.1) Despite subsection (1), no manufacturer shall use a preparation made by extending any of the following substances on a substratum of mica in a drug unless the requirements in subsection (4.3) are met:

    • (a) titanium dioxide (C.I. No. 77891);

    • (b) iron oxides (C.I. Nos. 77489, 77491, 77492, 77499); or

    • (c) a mixed oxide made from substances referred to in both paragraphs (a) and (b).

  • (4.2) Despite subsection (2), no person shall import for sale a drug that contains a preparation made by extending any of the substances referred to in paragraphs (4.1)(a) to (c) on a substratum of mica unless the requirements in subsection (4.3) are met.

  • (4.3) The requirements referred to in subsections (4.1) and (4.2) are the following:

    • (a) the drug shall be in a solid dosage form intended for oral administration or in a liquid dosage form intended for oral administration or it shall be a drug intended for external use;

    • (b) in the case where the drug is in a solid dosage form intended for oral administration, the drug shall not contain more than 3% of the preparation;

    • (c) in the case where the preparation was made using iron oxides, the preparation shall not contain more than 55% iron oxides.

  • (5) Subsections (1), (2), (4.1) and (4.2) do not apply in respect of a drug that is represented as being solely for use in the disinfection, for disease prevention, of

    • (a) medical devices;

    • (b) health care facilities; or

    • (c) premises in which food is manufactured, prepared or kept.

  • SOR/84-949, s. 1
  • SOR/86-590, s. 1(E)
  • SOR/94-460, s. 1
  • SOR/95-431, s. 1
  • SOR/2002-369, s. 1
  • SOR/2005-95, s. 1
  • SOR/2018-248, s. 1
.

Prescription Drugs

 In deciding whether to amend the Prescription Drug List in respect of a drug, including by adding the drug to it or removing the drug from it, the Minister shall consider whether any of the following criteria apply with respect to the drug:

  • (a) supervision by a practitioner is necessary

    • (i) for the diagnosis, treatment, mitigation or prevention of a disease, disorder or abnormal physical state, or its symptoms, in respect of which the drug is recommended for use, or

    • (ii) to monitor a disease, disorder or abnormal physical state, or its symptoms, in respect of which the drug is recommended for use, or to monitor the use of the drug;

  • (b) the level of uncertainty respecting the drug, its use or its effects justifies supervision by a practitioner; or

  • (c) use of the drug can cause harm to human or animal health or a risk to public health and the harm or the risk can be mitigated by a practitioner’s supervision.

  • SOR/2013-122, s. 10
  • SOR/2017-18, s. 21(F)
.

 The Minister shall consult the general public on any proposal by the Minister to remove a drug from the Prescription Drug List.

  • SOR/2013-122, s. 10
.

 Sections C.01.040.3 and C.01.040.4 apply, with any modifications that the circumstances require, in respect of classes of drugs.

  • SOR/2013-122, s. 10
.
  •  (1) No person shall sell a prescription drug unless

    • (a) they are entitled under the laws of a province to dispense a prescription drug and they sell it in that province under a verbal or written prescription that they received; or

    • (b) they sell it under section C.01.043.

  • (2) In the case of a verbal prescription, the person referred to in paragraph (1)(a) or a pharmacy technician shall create a written record of the prescription that includes the following information:

    • (a) the date on which the prescription was received and, if applicable, the number of the prescription;

    • (b) the name and address of the person to whom the prescription was issued;

    • (c) the proper name, common name or brand name of the drug and its quantity;

    • (d) the person’s name and the name of the practitioner who issued the prescription; and

    • (e) the directions for use provided with the prescription, whether or not the practitioner authorized it to be refilled and, if refills are authorized, the number of authorized refills.

  • (3) The person referred to in paragraph (1)(a) shall retain the written prescription referred to in subsection (1) or the record referred to in subsection (2) for at least two years after the day on which the prescription is filled.

  • SOR/78-424, s. 2
  • SOR/80-543, s. 3
  • SOR/93-202, s. 6
  • SOR/93-407, s. 2
  • SOR/2013-122, s. 11
.

 Subject to paragraph C.01.041.3(2)(b), a pharmacist or pharmacy technician may transfer to another pharmacist or pharmacy technician a prescription for a prescription drug.

  • SOR/78-424, s. 3
  • SOR/2013-122, s. 11
.
  •  (1) Before a pharmacist sells a drug under a prescription that was transferred under section C.01.041.1, the pharmacist or a pharmacy technician shall

    • (a) create a written record of the name and address of the pharmacist or pharmacy technician who transferred the prescription and, if applicable, the number of authorized refills remaining and the date of the last refill; and

    • (b) obtain a copy of the written prescription or of the written record that was created under subsection C.01.041(2), as the case may be, or, in the case of a verbal transfer, create a written record that includes the information referred to in that subsection.

  • (2) The pharmacist shall retain the documents referred to in subsection (1) for at least two years after the day on which the prescription was filled.

  • SOR/78-424, s. 3
  • SOR/2013-122, s. 11
.
  •  (1) A pharmacist or a pharmacy technician who transfers a prescription under section C.01.041.1 shall indicate the date of transfer on the original of the written prescription or of the written record created under subsection C.01.041(2) or in a record kept under the name of the patient in question, as the case may be.

  • (2) When the pharmacist or pharmacy technician has transferred the prescription,

    • (a) the pharmacist shall not make any additional sales under the prescription; and

    • (b) the pharmacist or pharmacy technician shall not transfer the prescription to another pharmacist or pharmacy technician.

  • SOR/78-424, s. 3
  • SOR/2013-122, s. 11
.

 A person referred to in paragraph C.01.041(1)(a) shall not refill a prescription for a prescription drug unless authorized by the practitioner and, in the case of such an authorization, they shall not refill a prescription more than the number of times specified by the practitioner.

  • SOR/78-424, s. 4
  • SOR/2013-122, s. 11
.

 A person referred to in paragraph C.01.041(1)(a) shall indicate on the original of or on the copy of the written prescription or the written record created under subsection C.01.041(2) or in a record kept under the name of the patient in question, as the case may be,

  • (a) the date on which the prescription was filled;

  • (b) the date of each refill, if applicable;

  • (c) the quantity of drug sold when the prescription was filled and, if applicable, for each refill; and

  • (d) the name of the person who sold the drug.

  • SOR/2013-122, s. 11
.
Loading…